
Water quality and hardness
This module explores environmental chemistry through four interconnected themes:
- Water hardness and treatment: Begin with Water quality and hardness, learn Expressing hardness as calcium carbonate and Worked problems on water hardness, then explore Ion-exchange softening and demineralization and Reverse osmosis and desalination.
- Potable water disinfection: Move to Water purification, chlorination and breakpoint and Ozone and ultraviolet disinfection.
- Wastewater and biological treatment: Examine Water pollution, dissolved oxygen, BOD and COD, Sewage treatment: stages and activated sludge, and Trickling filters and the UASB process.
- Global environmental sustainability: Conclude with E-waste and resource recovery, Greenhouse gases and climate change, Stratospheric ozone and ozone depletion, Sustainable development and the 17 SDGs, and Revision sheet and practice problems.
Water characteristics
Water is an effective solvent for many ionic and polar substances used across domestic households, agriculture, and industry. In industrial plants, large volumes of water serve as process solvents, reaction media, and heat-transfer fluids for cooling and steam generation. Because natural water dissolves minerals from soil and rock, its suitability depends directly on the dissolved ionic species present.
Hardness
Hardness is the characteristic of water that prevents rapid lathering with soap. It is primarily caused by dissolved multivalent metallic cations—principally calcium () and magnesium (), along with smaller amounts of other divalent or heavy metal ions such as and .
Hard water
Hard water contains appreciable concentrations of dissolved and salts (, , , , , and ). It does not lather readily with soap, instead forming a sticky white precipitate.
Soft water
Soft water contains negligible concentrations of and ions and produces a rich, persistent lather immediately upon agitation with soap.
Mechanism of soap scum formation
Commercial soaps consist of sodium or potassium salts of long-chain fatty acids, such as sodium stearate (). In soft water, sodium stearate dissolves freely to lower surface tension and assemble into cleansing micelles.
When soap is introduced to hard water, the dissolved and ions displace to form insoluble calcium stearate or magnesium stearate precipitates, collectively known as soap scum:
Both and precipitate as sticky, greyish-white curds that adhere to fabrics, piping, and washbasins. The hardness cations consume soap stoichiometrically before a persistent lather can develop, substantially increasing soap consumption.
Classification of water hardness
Total hardness is classified into two distinct forms based on the counter-anions present and their behaviour upon heating:
Temporary hardness (carbonate hardness)
Temporary hardness is caused by dissolved bicarbonates of calcium and magnesium, namely and . It is designated as carbonate hardness because it represents the fraction of hardness chemically equivalent to the bicarbonate and carbonate alkalinity in the water. Boiling expels dissolved carbon dioxide gas and shifts the equilibrium, precipitating insoluble calcium carbonate and, upon complete thermal decomposition, magnesium hydroxide (which possesses a much lower solubility product than magnesium carbonate):
Permanent hardness (non-carbonate hardness)
Permanent hardness is caused by dissolved chlorides and sulfates of calcium and magnesium (, , , and ). Because these neutral salts do not decompose into insoluble precipitates upon boiling, permanent hardness cannot be removed thermally and requires chemical softening (such as the lime-soda process), ion-exchange demineralization, or membrane separation.
Comparison of hardness types
| Feature | Temporary (Carbonate) Hardness | Permanent (Non-Carbonate) Hardness |
|---|---|---|
| Causative salts | , | , , , |
| Effect of boiling | Precipitates as and | Not removed by simple boiling |
| Removal methods | Thermal boiling or stoichiometric lime addition | Ion exchange, reverse osmosis, or lime-soda softening |
Operational impacts of hard water
Utilizing untreated hard water introduces significant technical difficulties across municipal, commercial, and industrial facilities:
- Domestic laundering and personal care: Soap is consumed wastefully by precipitating and before cleansing action begins, leaving curdy scum residues on skin, garments, and fixtures.
- Textile and sugar processing: In textile wet processing and dyeing, multivalent cations precipitate dyestuffs, causing shade variations and patchy spots. In sugar refining, dissolved and retard and inhibit the crystallization of sucrose.
- Boilers and heat-transfer equipment: When hard water is heated in high-pressure steam boilers, bicarbonates and sulfates deposit as dense, adherent mineral scales (, ) or loose sludges. Because mineral scale has an extremely low thermal conductivity, it acts as an insulator, drastically lowering thermal efficiency, elevating fuel consumption, and inducing severe localized tube overheating and rupture. Additionally, dissolved undergoes high-temperature hydrolysis:
The liberated hydrochloric acid () attacks boiler steel, causing severe localized pitting corrosion.
Watch out. Hard water is not inherently toxic or unsafe to drink, nor does it universally prohibit concrete mixing; in fact, moderate calcium and magnesium levels contribute to dietary mineral intake. Conversely, "soft water" does not mean pure or microbiologically safe water—sodium-cycle softened water still contains dissolved salts, and any microbial contamination needs appropriate disinfection.
Watch out. Do not confuse hardness (concentration of multivalent cations like and ) with alkalinity (acid-neutralizing capacity due to , , and ) or salinity (total dissolved salts). Sodium salts such as (salinity) and (alkalinity) do not contain divalent cations and therefore do not produce hardness.
Self-check questions
-
Question: Why does a water sample containing of dissolved lather easily with dilute soap solution, whereas a sample containing of forms a sticky precipitate? Answer: Sodium chloride () shares the cation with sodium stearate soap and does not precipitate fatty acid anions, whereas from reacts with stearate anions to precipitate insoluble calcium stearate scum.
-
Question: Which solid precipitates form when water containing both and is boiled thoroughly? Answer: Insoluble calcium carbonate () and magnesium hydroxide ().
Expressing hardness as calcium carbonate
Why Calcium Carbonate is the Universal Reference
Natural water contains a mixture of dissolved calcium and magnesium salts—such as bicarbonates, sulfates, and chlorides—each having a distinct molar mass. Directly summing their mass concentrations in is chemically meaningless because of contains a different number of divalent cations than of . To compare and sum these contributions on a uniform stoichiometric basis, water analysis expresses all hardness and alkalinity concentrations in terms of equivalents of calcium carbonate ().
Calcium carbonate is chosen as the universal reference standard for three key reasons:
- Convenient arithmetic: Its molecular mass is (using standard rounded atomic masses) and its equivalent mass is , which greatly simplifies stoichiometric conversions.
- Relevant chemistry: is the primary insoluble precipitate formed during scale formation and standard precipitation softening processes.
- Primary analytical standard: Highly pure, anhydrous can be directly weighed and dissolved in dilute hydrochloric acid to prepare primary standard calcium solutions for calibrating EDTA titrations.
Converting Concentrations to Equivalents
Because chemical reactions occur on an equivalent-to-equivalent basis, the number of equivalents of any hardness-producing species equals the number of equivalents of it represents:
For a chemical species present at mass concentration (in ) with equivalent mass (in or ), the hardness expressed in is:
Equivalent Mass
The equivalent mass of an ion or salt is its molar mass divided by the total positive (or negative) charge magnitude exchanged per formula unit (). For a salt or ion containing a single divalent cation (), :
Watch out. The shortcut applies only when one formula unit supplies exactly one divalent cation (, e.g., , , , ). For monovalent ions such as (, ) or multivalent salts such as (, ), you must evaluate the fundamental formula .
Equivalent Masses of Common Water Constituents
Using standard rounded atomic masses (, , , , , , , ):
| Species | Formula / Atomic Mass (, ) | Valence Factor (, ) | Equivalent Mass (, ) | Role in Water Analysis |
|---|---|---|---|---|
| Reference standard | ||||
| Non-carbonate hardness | ||||
| Non-carbonate hardness | ||||
| Non-carbonate hardness | ||||
| Non-carbonate hardness | ||||
| Carbonate hardness & alkalinity | ||||
| Carbonate hardness & alkalinity | ||||
| Total hardness cation | ||||
| Total hardness cation | ||||
| Bicarbonate alkalinity | ||||
| Alkalinity only (no hardness contribution) |
Units of Hardness and Concentration Conversions
Hardness is reported across engineering disciplines in several interconvertible units:
| Unit | Symbol | Definition | Relationship to |
|---|---|---|---|
| Milligrams per litre | Milligrams of equivalent per litre of water | ||
| Parts per million | Parts of equivalent per parts of water by mass | (for ) | |
| Milliequivalents per litre | Milliequivalents of hardness cations per litre of water | ||
| Degree French | Parts of equivalent per parts of water | ||
| Clarke's Degree | Grains of equivalent per Imperial gallon () |
Conversion from Molarity and Normality
When the concentration of a dissolved species is provided in chemical concentration units, use dimensional analysis:
-
Relation between Normality and Molarity:
-
Conversion of Normality to Equivalent:
Watch out. Normality is a reaction-specific concentration unit expressed in , not a dimensionless mass fraction like . Always verify that your final answer specifies units of volume concentration (), not pure mass ().
See Worked problems on water hardness for complete multi-ion numerical examples.
Hardness Classifications and Components
When comprehensive water analyses report individual ionic species rather than salt pairs, hardness components are resolved by comparing Total Hardness () with Total Alkalinity () (both expressed in ).
Total Hardness ()
Total Hardness is the stoichiometric sum of all divalent calcium and magnesium ions:
Total Alkalinity ()
Total Alkalinity in natural circumneutral waters is predominantly governed by the bicarbonate ion ():
Carbonate Hardness ()
Carbonate Hardness, also known as temporary hardness, is the portion of divalent cations chemically paired with carbonate or bicarbonate anions. Because carbonate hardness cannot exceed the total divalent cations available:
Non-Carbonate Hardness ()
Non-Carbonate Hardness, also known as permanent hardness, represents cations paired with non-alkaline anions such as , , or :
Alkalinity and Hardness Interactions
- When : All divalent hardness cations are paired with bicarbonate/carbonate ions. Under this condition, and . The excess alkalinity represents bicarbonate paired with monovalent alkali metal cations (such as ).
- Effect of Adding Sodium Salts: Adding to hard water containing does not alter (since causes no hardness), but it raises , converting non-carbonate hardness into carbonate hardness. In contrast, neutral salts like change neither nor .
- Practical Note on Boiling: Carbonate hardness is termed "temporary" because boiling precipitates calcium as . However, this is an operational classification; residual hardness persists due to the finite thermodynamic solubility product () of and .
Self-Check Questions
-
Question: A water sample displays and bicarbonate alkalinity (both as ). Determine the carbonate hardness () and non-carbonate hardness ().
- Answer: Since , . The non-carbonate hardness is . The surplus of alkalinity exists as non-hardness bicarbonates (e.g., ).
-
Question: Why is the equivalent mass of the bicarbonate ion taken as when calculating equivalents, given that of reacts with of ?
- Answer: Equivalent mass is defined strictly on a per-unit-charge basis (). Because carries a net charge of , its equivalent mass equals its formula mass (). Consequently, two equivalents of bicarbonate () stoichiometrically counterbalance one equivalent of divalent calcium (, representing or of ).
Worked problems on water hardness
Formulas and reference data
To compare waters containing different dissolved minerals, every hardness-producing species—and every alkalinity-producing species used to partition carbonate (temporary) and non-carbonate (permanent) hardness—is converted into its chemical equivalent of calcium carbonate (). Because the molar mass of is and its ionic charge factor , the equivalent mass of is .
For any dissolved salt or ion with mass concentration (in or ) and equivalent mass (in ):
When solution concentrations are reported in normality () or molarity (), where and is the total positive charge per formula unit:
| Species | Formula | Atomic / Molar mass () | Charge factor () | Equivalent mass (, ) |
|---|---|---|---|---|
| Calcium carbonate | ||||
| Calcium bicarbonate | ||||
| Magnesium bicarbonate | ||||
| Calcium sulfate | ||||
| Magnesium sulfate | ||||
| Calcium chloride | ||||
| Magnesium chloride | ||||
| Sodium bicarbonate | ||||
| Calcium ion | ||||
| Magnesium ion | ||||
| Bicarbonate ion |
Partitioning hardness
Total hardness ()
Total hardness is the sum of the equivalents of all multivalent hardness cations ( and salts or ions). Alkali metal salts (, ) do not contribute to .
Alkalinity ()
Alkalinity is the sum of the equivalents of all bicarbonate and carbonate species present, including sodium bicarbonate ().
Carbonate hardness ()
Carbonate hardness (temporary hardness) is defined by . If alkalinity is less than total hardness, . If alkalinity equals or exceeds total hardness, .
Non-carbonate hardness ()
Non-carbonate hardness (permanent hardness) is defined by . Because can never exceed , permanent hardness is never negative.
Watch out. Do not round individual salt equivalents to two decimal places and then add them. Truncating intermediate values (for example, writing instead of for ) introduces cumulative rounding errors in the final total. Keep at least four decimal places during intermediate steps.
Worked examples
Example 1: Single permanent hardness salt ()
Problem:
A water sample contains of per litre. Calculate its hardness in terms of equivalence.
Solution:
From the reference table, the molar mass of is and its equivalent mass is . Given :
Because is a sulfate salt, this entire is permanent (non-carbonate) hardness.
Example 2: Single temporary hardness salt ()
Problem:
Calculate the hardness of a water sample containing of calcium bicarbonate, .
Solution:
The molar mass of is and its equivalent mass is . Converting into equivalence:
Example 3: Mixed bicarbonate and sulfate salts
Problem:
Calculate the temporary (carbonate), permanent (non-carbonate), and total hardness of a water sample having the following composition: , , , and .
Solution:
Convert each hardness-producing salt to its equivalent using unrounded fractions first:
- ():
- ():
- ():
- ():
Sum into carbonate (temporary) and non-carbonate (permanent) contributions:
Example 4: Mixed bicarbonate, sulfate, and chloride salts
Problem:
Calculate the temporary, permanent, and total hardness of water containing , , , and .
Solution:
Calculate the equivalent of each salt:
- ():
- ():
- ():
- ():
Sum the respective components:
Example 5: Effect of sodium bicarbonate () on hardness partitioning
Problem:
Calculate the temporary, permanent, and total hardness of a water sample containing , , , , and .
Solution:
Sodium ions () do not precipitate soap or form boiler scale, so adds zero total hardness. However, its bicarbonate ion () increases the total alkalinity of the water. Upon boiling, this extra reacts with or ions originally paired with , precipitating them as carbonate scale. Thus, converts an equivalent amount of permanent hardness into temporary hardness while leaving unchanged.
From Example 3, the four and salts give:
- Bicarbonate alkalinity from and
Determine the alkalinity contribution of ():
Since , temporary (carbonate) hardness equals total alkalinity:
Example 6: Second mixture containing sodium bicarbonate ()
Problem:
Calculate the temporary, permanent, and total hardness of water having the composition: , , , , and .
Solution:
Convert the four divalent hardness salts to equivalents:
- ():
- ():
- ():
- ():
Include the equivalent of () in the alkalinity sum:
Example 7: Mixture containing neutral sodium chloride ()
Problem:
Calculate the temporary, permanent, and total hardness of water containing , , , , and .
Solution:
Watch out. Unlike , sodium chloride () contains neither a multivalent hardness cation (, ) nor a carbonate/bicarbonate alkalinity anion (, ). Therefore, has no effect on temporary, permanent, or total hardness. Never subtract or add an equivalent to permanent or temporary hardness.
Calculate the equivalents of the four hardness-producing salts:
- ():
- ():
- ():
- ():
- : Ignored ( contribution to both hardness and alkalinity).
Sum the carbonate and non-carbonate contributions:
Example 8: Ion-basis water analysis where
Problem:
Calculate the temporary, permanent, and total hardness of a water sample whose ionic analysis reports , , and (these are the relevant entries of a partial ion analysis).
Solution:
When an analysis is given in terms of individual ions rather than combined salts, total hardness depends solely on the hardness cations ( and ), while temporary hardness is governed by the bicarbonate alkalinity (). Monovalent ions (, ) and unaligned sulfate figures from incomplete ion balances are not needed.
-
Total Hardness ():
- ():
- ():
-
Bicarbonate Alkalinity ():
- ():
- ():
-
Carbonate and Non-Carbonate Hardness:
Because :
Example 9: Ion-basis water analysis where alkalinity exceeds total hardness ()
Problem:
A sample of water on analysis gave the following results: , , , and . Calculate the temporary, permanent, and total hardness.
Solution:
-
Total Hardness ():
- ():
- ():
-
Bicarbonate Alkalinity ():
- ():
- ():
-
Partitioning and :
Here, the total bicarbonate alkalinity () exceeds the total hardness (); the reported ion list is incomplete and must not be treated as a charge-balanced full analysis. Sodium does not contribute to total hardness. Because temporary (carbonate) hardness can never exceed total hardness ():(Do not report temporary hardness as or permanent hardness as ; is the alkalinity, not the temporary hardness.)
Example 10: Hardness from normality and molarity of and solutions
Problem:
Calculate the hardness (in ) of:
- solution
- solution
Solution:
-
solution:
Using : -
solution:
Each formula unit of releases one ion (). First convert molarity to normality ():
Example 11: Formal charge equivalence of
Problem:
Calculate the formal equivalent concentration of a solution.
Solution:
One formula unit of dissociates into two cations, giving a total positive charge factor .
Important chemical caveat: In water quality practice, routine hardness is defined by and . Aluminium salts such as (alum) undergo rapid acid hydrolysis in water to form insoluble flocs during coagulation rather than remaining as simple dissolved hardness ions. Therefore, this calculation represents a formal stoichiometric charge equivalence used in textbook exercises, not a standard natural-water hardness measurement.
Self-check questions
-
Question: A water sample has and . What are its carbonate (temporary) hardness and non-carbonate (permanent) hardness?
Answer: Carbonate hardness , and non-carbonate hardness . -
Question: Why does adding of to a hard water sample leave both temporary and permanent hardness unchanged, whereas adding of can increase temporary hardness and decrease permanent hardness?
Answer: supplies neither hardness cations (, ) nor alkalinity anions (). By contrast, supplies alkalinity, which pairs with non-carbonate ions upon boiling, increasing the carbonate portion up to the total hardness; once all hardness is carbonate hardness, further bicarbonate cannot increase it.
Ion-exchange softening and demineralization
Ion exchange
Ion exchange is a reversible heterogeneous process in which mobile ions from an aqueous solution are exchanged for stoichiometrically equivalent, similarly charged ions electrostatically bound to an insoluble solid matrix. In water treatment, these solid matrices are synthetic organic polymers known as ion-exchange resins. They possess a high-molecular-weight, porous, three-dimensional cross-linked structure—most commonly a styrene–divinylbenzene copolymer—that prevents the resin from dissolving in water while allowing dissolved ions to diffuse freely through its micropores.
Ion-exchange resin
An ion-exchange resin consists of two distinct components:
- Fixed functional groups: Covalently bonded to the immobile polymer backbone, carrying a net charge when ionized. Strong-acid sulfonate and strong-base quaternary ammonium resins remain ionized across broad operational pH ranges.
- Mobile counter-ions: Oppositely charged ions loosely associated with the fixed groups that can reversibly exchange with ions of like charge in the surrounding aqueous medium.
Watch out. Student notes sometimes misspell the process as the "iron exchange process" or write resin formulas as and , which implies the entire resin carries a net charge. Always represent the neutral resin unit as a fixed polymer ion paired with a mobile counter-ion, such as (or simply ) for a cation exchanger and (or ) for an anion exchanger.
Softening versus demineralization
It is critical to distinguish between simple water softening and complete demineralization (deionization):
- Sodium-cycle cation softening: Employs a cation-exchange resin in the sodium form (). As hard water passes through the bed, divalent hardness cations ( and ) displace the monovalent ions on the resin: . Although hardness is removed, the effluent retains all dissolved anions and an equivalent amount of dissolved sodium salts (, ). Hence, the water is softened, not demineralized.
- Two-bed demineralization: Employs a hydrogen-form cation-exchange resin () placed in series before a hydroxide-form anion-exchange resin (). Cations are replaced by and anions are replaced by , which then combine to form pure water (), eliminating virtually all dissolved mineral electrolytes.
Resin classification
| Resin Type | Polymer Matrix & Functional Group | Fixed Group | Mobile Counter-Ion | Target Ions Removed |
|---|---|---|---|---|
| Cation Exchange Resin (CER) | Sulphonated or carboxylated styrene–divinylbenzene copolymer | Sulfonate () or carboxylate () | ||
| Anion Exchange Resin (AER) | Quaternary ammonium styrene–divinylbenzene or amine–formaldehyde copolymer | Quaternary ammonium () |
Cation exchange resins ()
Cation exchange resins contain acidic functional groups, synthesized by sulphonating a cross-linked styrene–divinylbenzene copolymer so that sulfonate groups () attach directly to the aromatic rings. In their hydrogen form, they release ions in stoichiometric exchange for metallic cations present in the raw water.
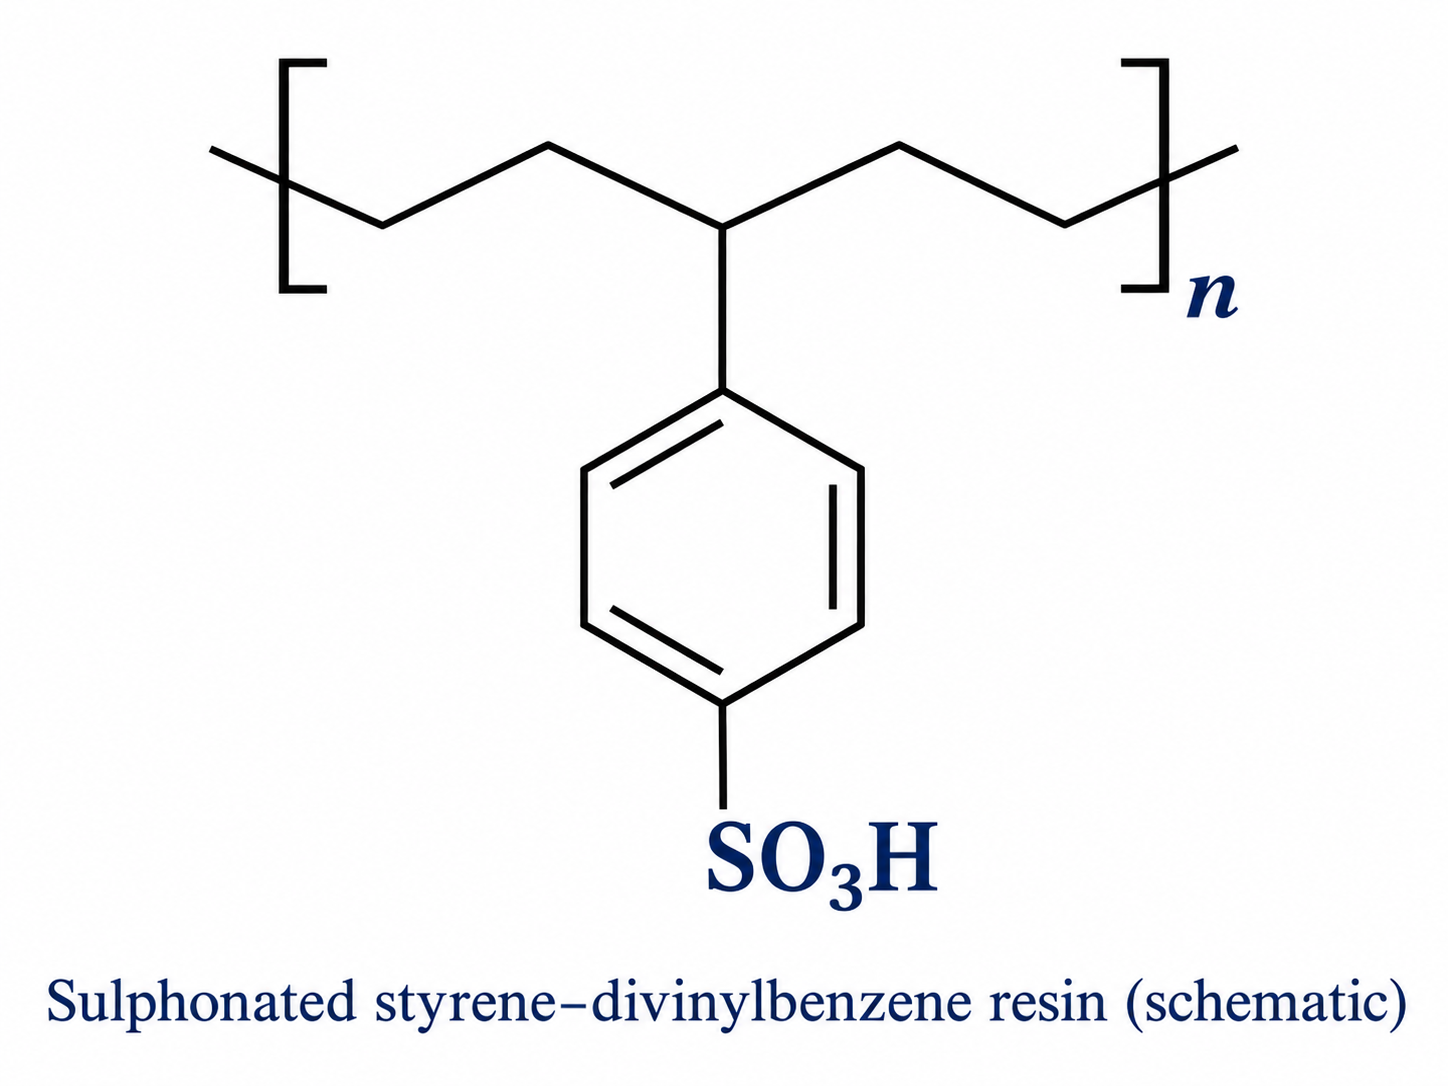
Figure 4.1: Structure of a sulphonated styrene–divinylbenzene cation-exchange resin with fixed groups and mobile counter-ions.
Anion exchange resins ()
Anion exchange resins contain basic functional groups, prepared by introducing quaternary ammonium groups () or tertiary amine groups onto the cross-linked copolymer matrix and treating the product with dilute . In their hydroxide form, they release ions in exchange for anions present in the incoming water.
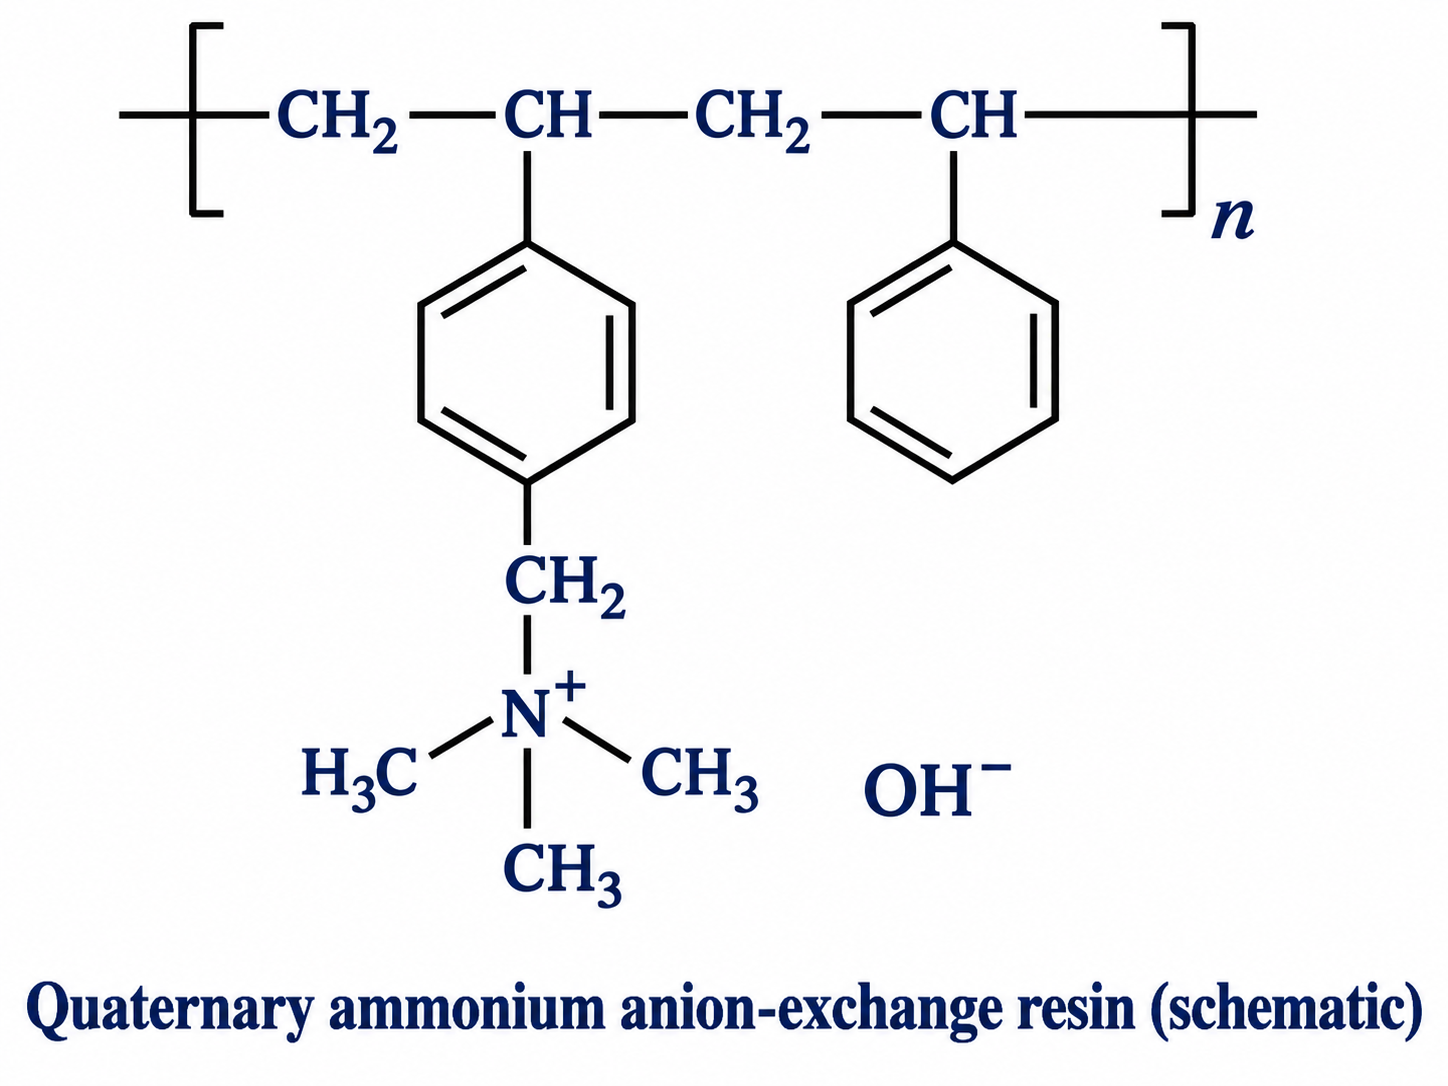
Figure 4.2: Structure of a quaternary ammonium styrene–divinylbenzene anion-exchange resin with fixed groups and mobile counter-ions.
Two-column demineralization process
Complete demineralization is achieved by routing raw feed water sequentially through two packed resin columns.
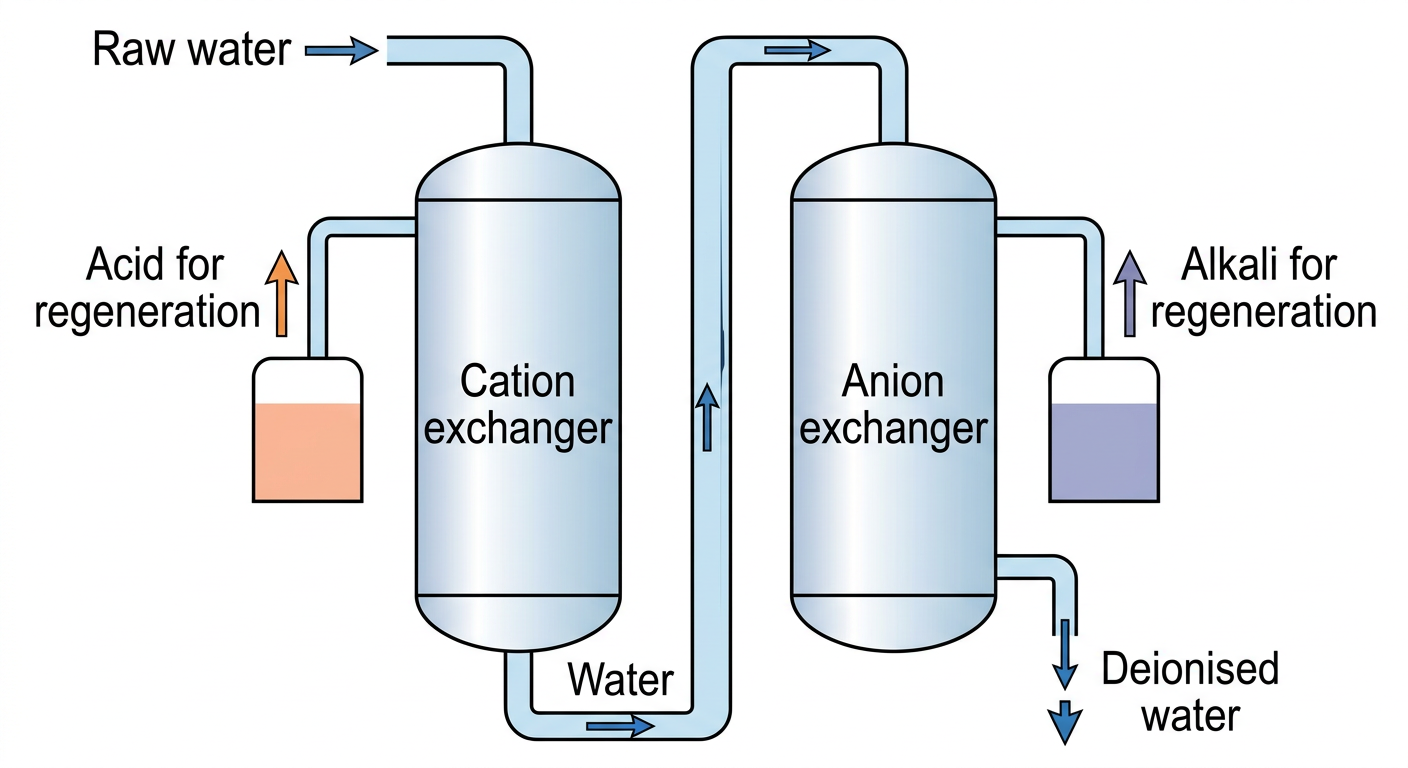
Figure 4.3: Two-stage ion-exchange demineralization unit showing the cation-exchange bed followed by the anion-exchange bed and their respective acid and alkali regeneration lines.
-
Passage through the cation-exchange column (CER): Raw water enters the column packed with resin. All dissolved metallic cations () are bound by the resin, displacing an equivalent quantity of ions into solution: and . When expressed for dissolved salts such as , the exchange generates free hydrochloric acid: . Similarly, dissolved sulphates form dilute sulphuric acid (). The effluent leaving the cation column is free of metallic cations but is distinctly acidic ().
-
Passage through the anion-exchange column (AER): The acidic effluent is fed into the second column containing resin. Dissolved anions () exchange with the hydroxyl counter-ions: and . Because the water entering this bed contains free mineral acids formed in the first step, the overall exchange directly neutralizes the acid molecules: and .
-
Neutralization and deionization: The ions released from the cation bed and the ions released from the anion bed combine instantaneously: . The effluent emerging from the second unit contains negligible dissolved ionic solutes when operated properly. This purified product is termed demineralized or deionized water.
Column sequencing
In standard two-bed systems, hard water must never be fed directly into the hydroxide-form anion exchanger (). If divalent cations () encounter high concentrations of ions prior to cation removal, insoluble metal hydroxides precipitate directly within the bed: . This precipitate coats the resin beads and clogs the interstitial voids of the column, causing severe pressure drop and halting ion exchange. Passing water through the acidic cation column first maintains all chemical species in soluble form throughout the process.
Regeneration of exhausted beds
With prolonged operation, the active counter-ions ( and ) become completely replaced by ions from the raw feed, rendering the beds exhausted. Because ion exchange is an equilibrium-driven process, passing concentrated acid or alkali solutions across the beds drives the equilibrium backward, restoring the active forms:
-
Regeneration of the cation exchanger: Flushing the exhausted bed with dilute hydrochloric acid () or dilute sulphuric acid () strips bound and ions from the polymer matrix: .
-
Regeneration of the anion exchanger: Flushing the exhausted bed with dilute sodium hydroxide () reloads the fixed sites with hydroxide counter-ions: and .
The soluble washings (, , ) are drained off as waste, and the columns are rinsed thoroughly with demineralized water before returning to service.
Merits and limitations
-
Advantages:
- Highly versatile; can treat varied feed compositions when paired with appropriate resin selections and pretreatment steps.
- Yields water with virtually zero residual hardness and exceptionally low electrical conductivity, meeting stringent feed-water specifications for high-pressure boilers and lead–acid battery electrolyte preparation.
- Produces no bulky sludge during routine service runs; resins can undergo repeated regeneration cycles over many years.
-
Limitations:
- High capital cost for specialized resin beds, column construction materials, and continuous consumption of regenerant chemicals ().
- Pretreatment requirement: Suspended solids, colloidal clay, oils, and multivalent transition metals () foul the resin pores, permanently blinding active exchange sites. Feed water must undergo prior coagulation and filtration to avoid resin degradation.
Self-check questions
-
Why is the effluent from the first column of a two-bed demineralizer acidic, and how is this acidity eliminated in the second column? Answer: In the cation-exchange column (), every dissolved cation () extracted from the water releases an equivalent number of ions, transforming salts like and into mineral acids ( and ). In the anion-exchange column (), the anions () are exchanged for ions, which neutralize the free ions directly to produce .
-
A student claims that passing hard water through a sodium-form cation resin () produces deionized water suitable for a high-pressure boiler. Critique this statement. Answer: The claim is incorrect. An resin exchanges and for ions (). While this softens the water by removing scale-forming divalent ions, the effluent retains an equivalent concentration of dissolved sodium salts (). Because these dissolved electrolytes contribute significant electrical conductivity and total dissolved solids (TDS), the water is softened but not demineralized.
Reverse osmosis and desalination
Desalination
Desalination is the process of removing excess dissolved salts and minerals from brackish water or seawater to produce fresh water suitable for domestic or industrial use. Unlike ion-exchange softening, which is uneconomical for high-salinity waters due to rapid resin exhaustion, membrane-based reverse osmosis (RO) is widely used to desalinate saline feedwater.
Osmosis and osmotic pressure
Natural osmosis
Natural osmosis is the spontaneous net movement of solvent from a dilute solution (higher water chemical potential) to a concentrated solution (lower water chemical potential) when two aqueous solutions of different solute concentrations are separated by a semipermeable membrane (which selectively allows water molecules to pass while rejecting most dissolved ions and larger molecules).
Osmotic pressure
Osmotic pressure (, typically measured in or ) is the hydrostatic pressure difference that builds up across the membrane at equilibrium—or the minimum excess pressure that must be applied to the concentrated side just to halt this net solvent flow. For dilute ideal solutions, osmotic pressure increases with molar concentration of the solute before dissociation and absolute temperature according to the van 't Hoff relation:
where:
- is the van 't Hoff factor
- is the molar concentration of solute
- is the universal gas constant
- is the absolute temperature
Principle of reverse osmosis
When external hydrostatic pressure is applied to the concentrated saline side, the direction of water transport depends on the applied transmembrane pressure difference () relative to the osmotic pressure difference () between the concentrated feed and dilute product streams:
- : Natural osmosis occurs; water flows spontaneously from the dilute side to the concentrated side.
- : Osmotic equilibrium is established; there is zero net flow of water across the membrane.
- : Reverse osmosis occurs; water is forced against the osmotic gradient from the concentrated saline feed into the dilute permeate stream.
Watch out. Do not compare osmotic pressure simply to absolute atmospheric pressure . Reverse osmosis requires the applied transmembrane pressure difference () to exceed the osmotic pressure difference () plus hydraulic friction and membrane resistance losses so that a practical permeate flux is achieved.
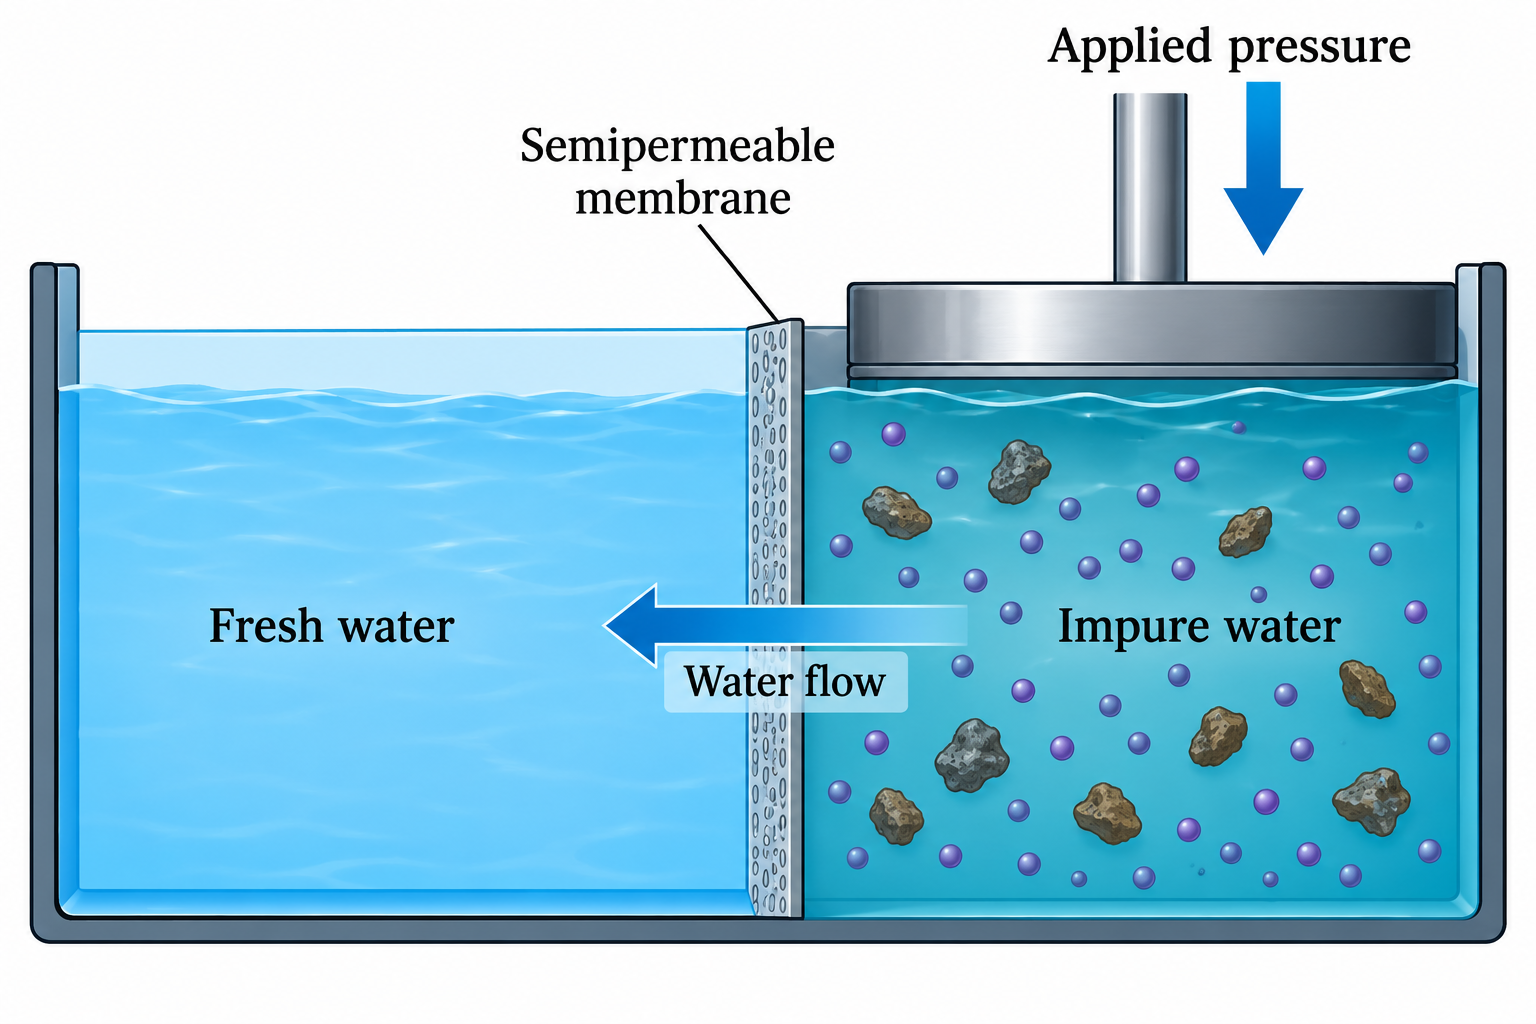
Figure: Principle of reverse osmosis—applying a pressure greater than the osmotic pressure forces water from the concentrated salt solution through a semipermeable membrane.
Process steps and membrane materials
An industrial RO desalination unit operates as a continuous cross-flow filtration system rather than a dead-end batch piston:
- Pretreatment: Raw seawater or brackish water is screened, coagulated, filtered, and chemically conditioned (e.g., with antiscalants and dechlorination agents) to remove suspended solids, biological foulants, and free chlorine that would foul or chemically degrade the membrane.
- Pressurization: A high-pressure feed pump raises the pretreated feedwater pressure well above its osmotic pressure (the exact value depends on feed salinity, membrane specifications, and targeted water recovery).
- Membrane separation: Pressurized feedwater enters the membrane module. Semipermeable membranes widely used in engineering practice include cellulose acetate and thin-film composite aromatic polyamide membranes. Water permeates through the dense active layer to form the low-salinity permeate (product stream), while rejected salts, organics, and colloids remain in the pressurized concentrate (reject or brine stream).
- Post-treatment and brine management: Permeate is stabilized (pH and remineralization) and disinfected for distribution, while the high-salinity concentrate is routed through energy-recovery devices and discharged or treated.
Advantages and limitations
Advantages
- Broad contaminant rejection: Rejects a high fraction of dissolved ionic salts alongside non-ionic contaminants such as dissolved organic matter, bacteria, and colloidal silica (which freely passes through conventional ion-exchange resin beds).
- No thermal phase change: Because water does not undergo vaporization, RO generally has a lower primary energy consumption than thermal desalination processes.
- Continuous operation: Eliminates periodic acid/alkali chemical regeneration steps required by ion-exchange beds, requiring only periodic cleaning-in-place (CIP) protocols.
Watch out. Reverse osmosis does not universally remove a fixed or of all ions, nor does it always have low capital cost. Rejection depends on membrane type, ion charge/size, feed salinity, and operating conditions, and high-pressure pumps, pretreatment systems, and membranes represent significant capital and electrical operating costs.
Limitations
- High electrical energy demand: High-pressure pumps consume substantial electrical energy, particularly for high-salinity seawater feeds.
- Membrane fouling and scaling: Suspended solids, biofouling, and precipitation of sparingly soluble salts (, , silica) deteriorate permeate flux and reduce membrane service life.
- Concentrate disposal: Safe discharge of large volumes of hyper-saline reject brine poses ecological challenges in inland or marine environments.
Comparison: Ion exchange vs. Reverse osmosis
| Feature | Ion-exchange demineralization | Reverse osmosis (RO) |
|---|---|---|
| Primary driving force | Chemical exchange of ions on functional resin sites | Applied transmembrane pressure () |
| Contaminants removed | Dissolved ionic species only | Dissolved ions, organics, colloids, and colloidal silica |
| Best suited feedwater | Low-to-moderate total dissolved solids (TDS) | Brackish water, seawater, or moderate-to-high TDS water |
| Main waste stream | Acidic and alkaline spent regeneration effluents | Continuous concentrated saline brine (reject) |
Self-check questions
-
Question: Why is reverse osmosis able to remove colloidal silica from boiler feedwater whereas an ion-exchange demineralizer cannot? Answer: Colloidal silica is non-ionic (uncharged polymeric particles) and therefore cannot participate in stoichiometric ion exchange on resin sites. An RO membrane rejects it mechanically and electro-dynamically via size exclusion and dense diffusion-barrier mechanisms.
-
Question: What happens in an RO cell if the applied transmembrane pressure equals the osmotic pressure difference across the membrane? Answer: The system reaches thermodynamic equilibrium (), resulting in zero net flux of solvent across the semipermeable membrane.
Water purification, chlorination and breakpoint
Natural waters contain suspended solids, colloids, dissolved minerals, organic impurities, and pathogenic microorganisms. Municipal water treatment employs a sequential multi-barrier train combining physical and chemical clarification to eliminate turbidity, followed by chemical disinfection to ensure microbiological safety before distribution.
Municipal water purification train
A conventional municipal water treatment plant operates through the following sequential unit operations:
- Screening: Raw intake water passes through coarse bar screens followed by fine mesh screens to trap floating debris (twigs, leaves, plastics, and rags) and protect downstream pumps from mechanical clogging or damage.
- Aeration: Water is intimately contacted with atmospheric air via spray nozzles, cascade steps, or diffused bubbles. Aeration expels dissolved volatile organic compounds and foul-smelling gases like , enriches dissolved oxygen, and oxidizes soluble ferrous () and manganous () ions into insoluble hydroxides for subsequent sedimentation.
- Coagulation and flocculation: Colloidal particles ( to ) remain indefinitely suspended due to mutual electrostatic repulsion from negative surface charges. Coagulant addition—typically alum, —supplies trivalent cations that compress the electrical double layer and neutralize surface charges. Hydrolysis of alum produces an insoluble, gelatinous aluminium hydroxide precipitate:
The liberated acid consumes natural alkalinity (predominantly ) to maintain the pH within the optimum precipitation window (). Slow, gentle mechanical mixing (flocculation) promotes particle collisions, aggregating micro-flocs into heavy, settleable flocs. 4. Sedimentation: Water enters large settling basins (clarifiers) under quiescent flow conditions with retention times of . Gravity pulls the heavy flocs and silt to the floor as clarifier sludge, removing of suspended solids. 5. Filtration: Clarified effluent percolates through graded sand and gravel beds (rapid gravity sand filters). This stage captures residual micro-flocs, colloidal silt, and protozoan cysts (such as Giardia and Cryptosporidium). While producing clear water, filtration alone cannot guarantee pathogen elimination, requiring downstream chemical disinfection. 6. Disinfection: The final chemical barrier destroys or permanently inactivates pathogenic bacteria, viruses, and amoebic cysts without necessarily achieving total sterilization. Chlorine, hypochlorites, chloramines, chlorine dioxide, and ultraviolet (UV) irradiation serve as primary municipal disinfectants.
Chemistry of chlorination
Chlorination remains the global standard for municipal disinfection because it is cost-effective, operates reliably over a broad temperature range, and leaves a persistent, measurable residual throughout distribution pipelines.
When elemental chlorine dissolves in water, it undergoes rapid disproportionation (hydrolysis) to form hypochlorous acid () and hydrochloric acid:
Hypochlorous acid is a weak acid ( at ) that partially ionizes into hypochlorite ions ():
Free Available Chlorine
Free available chlorine is the combined concentration of dissolved gaseous , unionized hypochlorous acid (), and hypochlorite ion ().
The disinfection efficacy of chlorine is strongly pH-dependent:
- Below : Unionized dominates. Because it is neutral and structurally compact, readily diffuses across the negatively charged cell walls and lipid membranes of bacteria. Inside the cell, it oxidizes vital enzymes and disrupts sulfhydryl () linkages.
- Above : The equilibrium shifts toward the negatively charged hypochlorite ion (). The negative bacterial membrane electrostatically repels , making it roughly times slower as a germicide than .
Watch out. Older textbooks often write and attribute disinfection to "nascent oxygen" (). That is an obsolete shorthand cartoon, not the real chemical mechanism. Hypochlorous acid () is the direct active oxidizing and chlorinating agent that damages microbial enzymes and cell membranes.
Hypochlorites and bleaching powder
To eliminate the safety and transport hazards of pressurized liquefied gas cylinders, smaller treatment works utilize sodium hypochlorite () solutions or solid calcium hypochlorite.
Bleaching Powder
Bleaching powder is a non-stoichiometric basic mixture of calcium hypochlorite , basic calcium chloride , and slaked lime , simplified empirically as or .
Upon dissolution in water, calcium hypochlorite dissociates to liberate hypochlorite ions, which hydrolyze to establish the active equilibrium:
Watch out. Do not write . Adding bleaching powder to neutral water yields aqueous calcium, chloride, and hypochlorite/hypochlorous acid species—the active disinfectant is described by the hypochlorite–hypochlorous acid equilibria.
Using bleaching powder carries notable operational drawbacks compared to gaseous or liquid :
- Instability: Atmospheric moisture and drive auto-decomposition during storage, degrading available chlorine content.
- Water hardening: The added and ions elevate total hardness and raise the water pH, shifting the equilibrium toward the less active .
- Sludge generation: Significant insoluble lime residues require frequent clarifier and dosing equipment cleanouts.
Chlorine demand and breakpoint chlorination
Chlorine Demand
Chlorine demand is the amount of applied chlorine consumed by reactions with oxidizable impurities before any persistent residual is established:
All terms are customarily expressed in or as .
Combined Available Chlorine
Combined available chlorine represents chloramines formed when hypochlorous acid reacts in competitive, stepwise substitution reactions with dissolved ammonia () or organic amines:
While monochloramine is a weaker disinfectant than free , it is substantially more stable, does not readily form trihalomethanes, and provides persistent secondary residual protection in expansive distribution mains.
Progressive addition of chlorine to water containing ammonia and reducing agents yields the classical breakpoint chlorination curve.
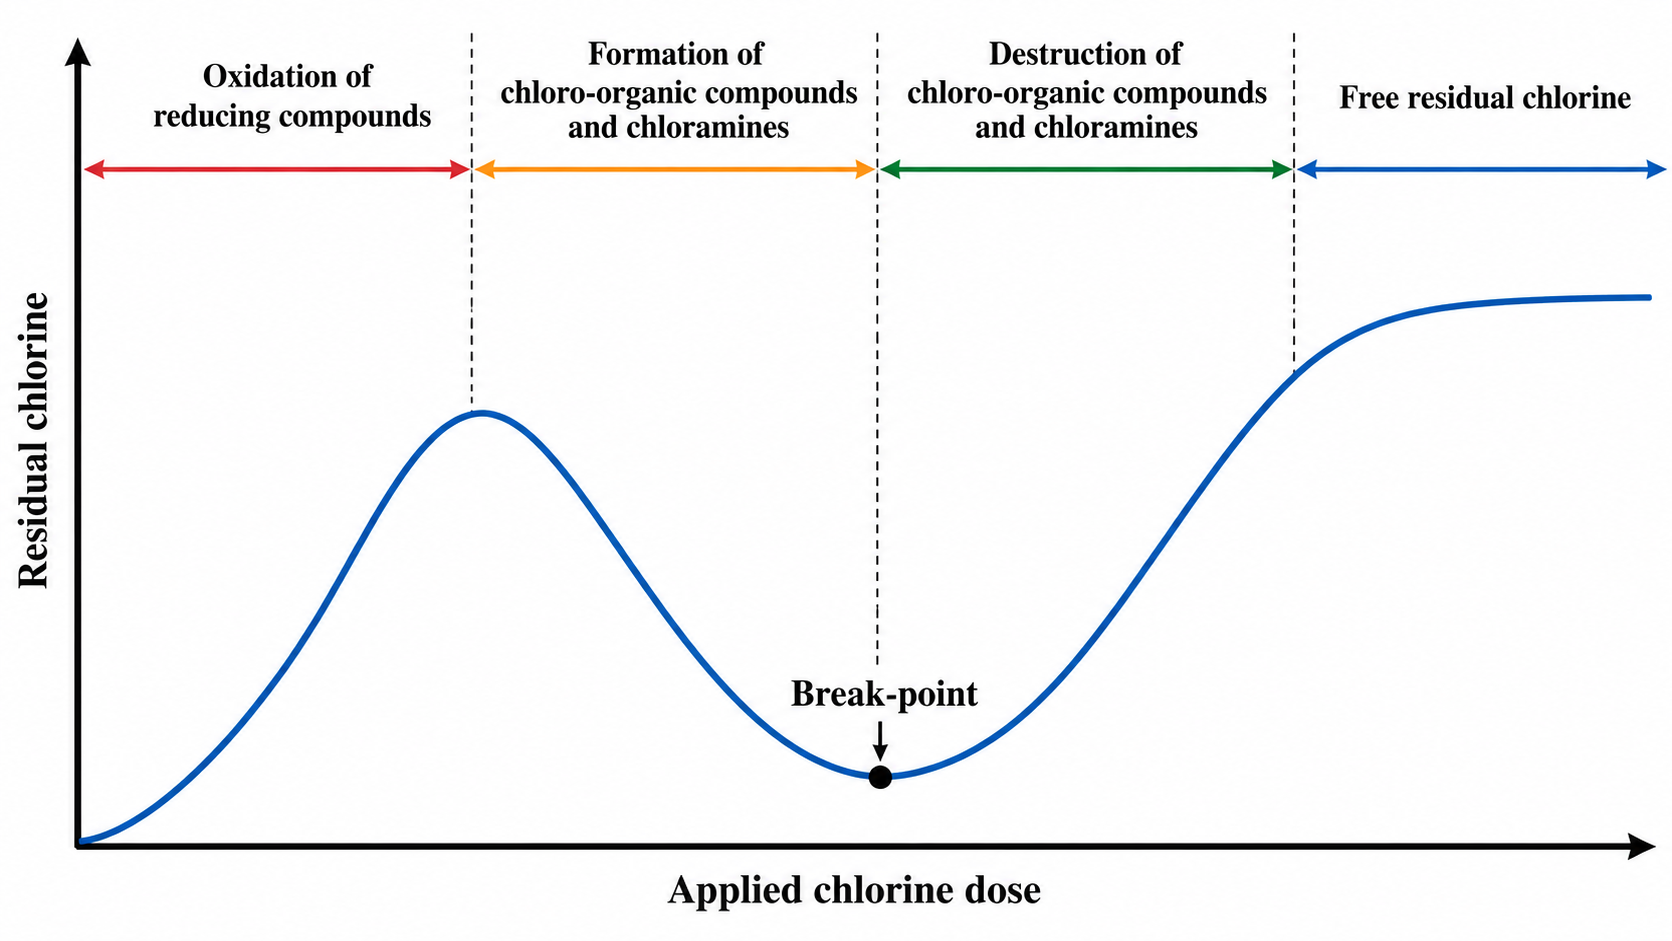
Figure: Breakpoint chlorination curve showing how measured chlorine residual changes with applied chlorine dose across four reaction stages.
| Reaction stage | Dominant chemical process | Residual behaviour |
|---|---|---|
| Stage I (initial demand) | Rapid stoichiometric reduction of chlorine to inactive by strong inorganic reducing agents (, , , ). | No measurable residual appears (residual = ). |
| Stage II (combined rise) | Reaction of with ammonia and organic amines to form chloramines (, ) and chloro-organics. | Total residual rises linearly to a local maximum; consists entirely of combined residual. |
| Stage III (chloramine destruction) | Excess oxidizes chloramines to volatile nitrogen species (, minor ) and inert chloride (). | Measured residual drops steeply to a distinct minimum value. |
| Stage IV (post-breakpoint) | Ammonia demand and readily oxidizable substances are completely consumed; additional chlorine remains unreacted as free and . | Residual rises linearly at a slope with further applied dose (free available chlorine). |
Breakpoint
Breakpoint is the exact dosage corresponding to the local minimum in residual chlorine at the end of Stage III.
Dosing past the breakpoint achieves:
- Destruction of chloramines responsible for foul "swimming pool" chlorinous odours and taste.
- Complete oxidation of ammonia compounds.
- Establishment of a reliable, fast-acting free residual chlorine baseline () to prevent recontamination in municipal mains.
Dechlorination and operational controls
Dechlorination
Dechlorination is the intentional reduction of excessive residual chlorine following disinfection contact time to avoid medicinal chlorophenol tastes, pipeline corrosion, or biological damage in downstream receiving waters.
Water is passed through granular activated carbon (GAC) filters or treated with chemical reducing agents such as sulphur dioxide (), sodium sulphite (), or sodium thiosulphate ():
Engineering parameters in municipal chlorination
- Contact time () and concentration (): Governed by Chick's Law and the empirical concept. Colder water reduces microbial inactivation kinetics, necessitating longer contact periods or higher disinfectant residuals.
- pH regulation: Optimal biocidal performance requires operating within to ensure that unionized predominates over .
- Turbidity shielding: Particulate matter protects shielded bacteria from chemical contact. Turbidity must be reduced below via upstream coagulation-filtration before chlorine addition.
- Disinfection by-product (DBP) mitigation: Excess free chlorine reacts with natural humic and fulvic organic matter to generate carcinogenic trihalomethanes (, e.g., ) and haloacetic acids (). Raw water must undergo efficient precursor removal before heavy terminal chlorination.
Self-check questions
-
Question: Why does the disinfecting efficiency of free chlorine drop significantly when the water pH rises from to ? Answer: Hypochlorous acid () has a . Below , unionized predominates, which readily crosses negatively charged microbial membranes to inactivate enzymes. Above , it dissociates into negatively charged hypochlorite ions (), which are electrostatically repelled by bacterial membranes, reducing biocidal efficiency by approximately two orders of magnitude.
-
Question: During breakpoint chlorination, why does the measured total chlorine residual decrease as more chlorine is added in Stage III? Answer: In Stage III, excess hypochlorous acid actively oxidizes the chloramines formed in Stage II into nitrogen gas (), nitrous oxide, and chloride ions (). Because nitrogen gas escapes and chloride has no disinfectant power, active combined chlorine is destroyed, driving the residual down to the breakpoint minimum.
Ozone and ultraviolet disinfection
Disinfection
Disinfection is the selective inactivation or destruction of disease-causing microorganisms (pathogens) to safe levels.
Sterilization
Sterilization refers to the complete destruction of all viable microbial life, including bacterial spores.
Municipal water treatment targets disinfection rather than full sterilization. While conventional chlorination remains widespread, physical and alternative chemical disinfectants—such as ozone () and ultraviolet (UV) radiation—effectively avoid undesirable chlorinous tastes, odours, and organohalogen by-products.
Ozone disinfection
Ozone () is an allotrope of oxygen and a powerful chemical oxidizing agent. Because ozone is thermodynamically unstable and decomposes rapidly, it cannot be stored or shipped in bulk and must be generated on-site.
On-site generation and delivery
Dry air or oxygen is passed through a silent electric discharge (corona discharge) across a high-voltage gap:
The resulting ozonized gas stream is introduced into water within a counter-current contact chamber to maximize gas-to-liquid mass transfer.
Disinfection mechanism
Unlike simplified textbook representations (), aqueous disinfection proceeds through two concurrent pathways:
- Direct oxidation: Molecular ozone () attacks specific functional groups, targeting proteins and bacterial cell walls.
- Indirect oxidation: Ozone reacts with water to form highly reactive, non-selective hydroxyl free radicals (), which rapidly lyse cell membranes and degrade nucleic acids.
Ozone also breaks down oxidizable organic molecules that produce colour, taste, and odour. However, it does not remove dissolved inorganic salts, hardness ions, or heavy metals.
Watch out. Do not refer to as "nascent "— is molecular dioxygen, and aqueous ozonation relies on dissolved and hydroxyl radicals (), not isolated atomic oxygen alone.
Merits and limitations
- Merits:
- Exceptional germicidal efficiency against bacteria, viruses, and chlorine-resistant cysts (Cryptosporidium, Giardia).
- Rapid reaction rate without imparting foreign chemical tastes or odours.
- Mineralizes oxidizable colour- and odour-causing organics.
- Limitations:
- High capital cost and substantial electrical energy consumption.
- In waters containing natural bromide (), ozone oxidizes bromide to form carcinogenic bromate ().
- Decomposes rapidly to molecular oxygen, leaving no residual disinfectant to protect distribution networks from recontamination.
Ultraviolet (UV) disinfection
Ultraviolet disinfection
Ultraviolet disinfection is a physical, chemical-free process that relies on germicidal light transmission through water. The germicidal window spans UV-C wavelengths (), with maximum biological effectiveness occurring near .
Operating principle
Water flows in thin profiles through an irradiation chamber housing low-pressure mercury lamps. The lamps are encased in protective quartz sleeves, which allow high transmission of UV-C light (unlike ordinary borosilicate glass, which absorbs shortwave UV).
The photons penetrate microbial cell membranes and are absorbed by cellular DNA and RNA. This photochemical excitation forms covalent cross-links—predominantly pyrimidine (thymine) dimers—between adjacent base pairs:
- Photon absorption disrupts the hydrogen bonding between complementary strands.
- The formed thymine dimers distort the nucleic acid helical backbone.
- DNA replication and transcription enzymes are arrested, preventing cellular reproduction.
Pathogens rendered unable to replicate cannot cause infection, successfully inactivating the microbes without chemically altering the bulk water.
Watch out. Conventional UV disinfection does not remove dissolved solids, hardness or heavy metals, and it only inactivates microbes when they receive a sufficient light dose.
Operational requirements
- Water clarity and transmittance: Suspended solids (turbidity) scatter and shield pathogens from light rays, while dissolved iron and humic substances absorb UV photons. Water must undergo coagulant-assisted filtration prior to UV treatment.
- Sleeve maintenance: Precipitation of hardness salts (, ) and biofilm accumulation on the quartz sleeve surfaces diminish radiation output; routine mechanical or chemical wiping is mandatory.
- Fluence control: System hydraulic flow rates must be regulated to ensure all fluid elements receive the required UV dose ().
- Absence of residual: Photons leave no active species in the treated water once it exits the reactor chamber.
Comparison of disinfection methods
| Feature | Chlorination () | Ozonation () | UV irradiation (UV-C, ) |
|---|---|---|---|
| Primary agent | Hypochlorous acid () | Dissolved and radicals | Germicidal photons (physical agent) |
| Mechanism | Chemical oxidation of cellular enzymes and structures | Chemical oxidation of cell walls and nucleic acids | Photochemical dimerization of DNA/RNA; halts replication |
| Distribution residual | Yes (free or combined residual protects distribution pipes) | None (rapidly decomposes to dissolved ) | None (acts strictly within the irradiation chamber) |
| Key merits | Low capital cost; reliable downstream pipe protection | Extremely fast pathogen kill; inactivates resistant cysts | Imparts no chemical tastes; produces no halogenated by-products |
| Key limitations | Generates DBPs (trihalomethanes, chlorophenols); chlorinous taste | High power cost; requires on-site generation; bromate formation risk | Ineffective in turbid water; requires clean quartz sleeves; zero residual |
Self-check questions
-
Why must water undergo filtration to remove turbidity before entering a UV disinfection unit? Answer: Suspended particles scatter, refract, and absorb UV-C photons, shielding embedded microorganisms from receiving the threshold radiation fluence required to dimerize their DNA.
-
Why do large municipal networks often add a small dose of chlorine after ozonation or UV treatment? Answer: Neither ozone nor UV leaves a persistent chemical residual in the water. A secondary booster dose of chlorine provides lasting downstream disinfection, preventing bacterial regrowth and neutralizing contaminants introduced through distribution pipe leaks.
Water pollution, dissolved oxygen, BOD and COD
Sources and Types of Water Pollution
Water Pollution
Water pollution occurs when undesirable foreign substances enter water bodies and degrade their physical, chemical, or biological quality, making them hazardous to aquatic ecosystems or human use.
Major sources fall into four primary categories:
- Domestic sewage: Municipal wastewater carries suspended solids, biodegradable organic waste, household detergents, and waterborne pathogens (enteric bacteria, viruses, and parasitic protozoa).
- Industrial effluents: Untreated or inadequately treated industrial discharges introduce synthetic organics (solvents, dyes, phenols), mineral acids, alkalis, and toxic heavy metals (such as , , , and ). Properly treated effluents eliminate or neutralize these contaminants prior to discharge.
- Agricultural runoff: Precipitation washing over cultivated fields transports dissolved nutrients (nitrates and phosphates from commercial fertilizers) and persistent agrochemicals (pesticides, herbicides) into surface waters. Excess nutrient loading drives eutrophication—rapid algal proliferation followed by algal die-off and severe oxygen depletion.
- Thermal discharges: Heated cooling water returned from thermal power stations and manufacturing plants raises the temperature of receiving waters, directly reducing oxygen solubility and accelerating the metabolic rates of aquatic organisms.
Aerobic vs. Anaerobic Decomposition
Microorganisms mineralize organic pollutants in water via two distinct biochemical pathways depending on the presence of dissolved molecular oxygen:
| Feature | Aerobic Decomposition | Anaerobic Decomposition |
|---|---|---|
| Oxygen requirement | Requires free, dissolved molecular oxygen (). | Occurs in the strict absence of dissolved molecular oxygen (). |
| Microorganisms | Driven by aerobic bacteria and fungi. | Driven by obligate and facultative anaerobes, including methanogenic archaea. |
| Typical end products | Stable, oxidized species: , , , , , and cellular biomass. | Reduced species: methane (), , organic acids, , , and cellular biomass. |
| Odour | Relatively odourless when adequately aerated. | Malodorous due to volatile reduced compounds (, mercaptans, ). |
| Energy & fuel value | High energy yield per mole of substrate; produces no fuel gases. | Low energy yield per mole of substrate; produces energy-rich biogas (). |
Using glucose () as a standard carbohydrate model, the degradation pathways are represented as:
-
Aerobic oxidation:
-
Anaerobic fermentation and methanogenesis:
Watch out. End products depend heavily on the wastewater substrate and microbial consortia present. Complete biological nitrification yields nitrate (), though nitrite () can form as an intermediate. Under anaerobic conditions, sulphate-reducing bacteria utilize sulphates as terminal electron acceptors, releasing toxic, foul-smelling hydrogen sulphide ().
Dissolved Oxygen (DO)
Dissolved oxygen (DO) is the concentration of uncompounded molecular oxygen gas () physically dissolved in an aqueous solution, expressed in milligrams per litre () or parts per million ().
DO does not refer to the chemically bonded oxygen in the molecule. It is crucial for maintaining higher aquatic life forms (such as fish and macroinvertebrates) and sustaining aerobic self-purification processes. DO is quantified analytically via Winkler’s iodometric titration or measured directly on-site using polarographic, galvanic, or optical luminescent DO probes.
Factors Governing Dissolved Oxygen Levels
- Temperature: Gas dissolution is an exothermic process. As water temperature rises, gas solubility declines; cold winter streams naturally retain substantially higher DO saturation levels than warm summer waters.
- Atmospheric pressure and re-aeration: In accordance with Henry's law, gas solubility is directly proportional to partial pressure. High-altitude waters experience lower barometric pressure and retain less DO at saturation. Mechanical turbulence, rapids, and artificial aeration increase the interfacial mass-transfer rate of atmospheric into the water.
- Salinity: Electrolytes decrease the solubility of non-polar gases through the "salting-out" effect; hence, seawater holds roughly less DO at saturation than freshwater under identical temperature and pressure conditions.
- Biological oxygen demand: Inflow of biodegradable organic matter triggers rapid aerobic microbial growth. When bacteria consume oxygen faster than natural re-aeration can supply it, the DO level drops rapidly toward hypoxia or anoxia.
Biochemical Oxygen Demand ()
Biochemical Oxygen Demand (BOD) is the mass of dissolved molecular oxygen required by aerobic microorganisms to biochemically oxidize the biodegradable organic matter (and oxidizable inorganic nitrogen, unless inhibited) present in one litre of wastewater under standardized laboratory incubation conditions: in complete darkness for ().
- Darkness ensures photosynthetic organisms (e.g., green algae) cannot generate in-situ oxygen during the test period.
- measures biodegradable organic load over an empirical window—it is neither a direct microbial population count nor a measure of total theoretical oxidation.
- A high signifies heavy biodegradable organic pollution capable of rapidly deoxygenating the receiving water body.
Worked Example: Calculating from a Diluted Sewage Sample
Problem: Raw sewage contains far more organic matter than the of oxygen available in saturated water, so the sample must be diluted before incubation. A aliquot of municipal wastewater is diluted to in a standard BOD bottle using oxygen-saturated, unseeded dilution water. The initial dissolved oxygen concentration () is . After of incubation at , the final dissolved oxygen concentration () is . Calculate the of the wastewater.
Assumptions: Dilution water is unseeded (zero seed blank depletion), and nitrification is uninhibited.
Solution:
-
Calculate the volumetric dilution fraction ():
-
Calculate for an unseeded sample:
-
Substitute the measured concentrations:
Chemical Oxygen Demand (COD)
Chemical Oxygen Demand (COD) is the mass of oxygen equivalent to the amount of a strong chemical oxidant consumed during the chemical oxidation of oxidizable organic and inorganic constituents in one litre of water under boiling, concentrated acidic reflux conditions (typically ).
- Oxidant system: Standardized potassium dichromate () in concentrated sulphuric acid (), refluxed in the presence of a silver sulphate () catalyst.
- Mechanism & titration: Orange dichromate ions () are reduced to green chromic ions (). The residual unreduced dichromate is back-titrated against ferrous ammonium sulphate (FAS), and the net oxidant consumed is converted stoichiometrically into equivalent .
- Chloride interference: Dissolved chloride ions () interfere by reacting with dichromate to generate chlorine gas (), causing an erroneously high COD reading: This interference is masked prior to digestion by adding mercuric sulphate (), which precipitates or complexes chloride as soluble, unreactive .
Watch out. For any given wastewater, . Acidic dichromate aggressively oxidizes both biologically degradable substrates and biologically recalcitrant organics (e.g., lignin, cellulose, synthetic dyes), along with oxidizable inorganics (, , ). Because chemical oxidation pathways differ fundamentally from microbial metabolic pathways, there is no universal conversion factor between BOD and COD; site-specific empirical ratios must be determined experimentally.
Comparison of DO, , and COD
| Parameter | What It Measures | Standard Units | Test Duration | Environmental Significance |
|---|---|---|---|---|
| DO | Free molecular dissolved in water at sampling time. | () | Immediate () | Assesses immediate capacity of water to support aquatic life. (High DO does not verify potability, as it does not detect pathogens or heavy metals). |
| Oxygen consumed biologically by microorganisms over at . | () | Measures biodegradable organic load and predicts downstream dissolved oxygen depletion in receiving waters. | ||
| COD | Oxygen equivalent of chemical oxidant () consumed under acid reflux. | () | Provides a rapid, comprehensive measure of total chemically oxidizable matter (biodegradable + non-biodegradable). |
Self-Check Questions
-
Why does a water sample from a fast-flowing mountain stream at low altitude on a winter morning have a higher DO than a stagnant pond on a hot summer afternoon? Answer: Three synergistic physical factors explain this:
- Temperature: Oxygen solubility is higher in cold winter water than in warm summer water.
- Atmospheric pressure: Lower elevation increases ambient barometric pressure, which raises equilibrium gas solubility via Henry's law.
- Aeration kinetics: Rapid turbulent flow constantly breaks the surface film, maximizing the rate of atmospheric re-aeration compared to stagnant pond water.
-
An industrial effluent containing synthetic dyes and cellulose shows and . Why is the COD so much larger than the ? Answer: The test only captures organic fractions readily metabolized by standard microbial cultures within . In contrast, the boiling digestion oxidizes virtually all organic species, including biologically recalcitrant aromatic dye rings and complex cellulose fibres, resulting in a substantially higher oxygen equivalent consumption.
Sewage treatment: stages and activated sludge
Sewage
Sewage refers to domestic and municipal wastewater generated by households, commercial establishments, and institutions. It carries human excreta, wash water, food waste, detergents, and pathogenic microorganisms.
Industrial Effluent
Industrial effluent originates from industrial processing plants and frequently contains heavy metals, recalcitrant synthetic organics, extreme acid/alkali loads, and toxic compounds requiring specialized chemical pretreatment.
The primary objective of municipal sewage treatment is to systematically reduce Biochemical Oxygen Demand (BOD), suspended solids, excess nutrients, and pathogens prior to environmental discharge or water reuse, thereby preventing eutrophication, dissolved oxygen depletion, and waterborne disease transmission.
Overview of Treatment Stages
Municipal sewage treatment progresses through distinct physical, biological, and chemical unit operations:
(Note: Settleable solids captured across primary and secondary clarifiers are routed to Sludge Processing: Thickening Anaerobic Digestion Dewatering Disposal/Beneficial Use.)
| Stage | Core Mechanism | Primary Target Pollutants | Typical Unit Operations |
|---|---|---|---|
| Preliminary | Physical interception & gravity settling | Coarse floating matter, rags, plastics, gravel, sand | Bar screens, fine mesh screens, grit chambers |
| Primary | Gravitational sedimentation | Settleable suspended solids, floating oils, grease | Primary sedimentation tanks (clarifiers) |
| Secondary | Biological metabolism & bio-flocculation | Colloidal and dissolved biodegradable organics (BOD) | Activated sludge systems, trickling filters, UASB reactors, secondary clarifiers |
| Tertiary | Advanced chemical, physical, and biological treatment | Plant nutrients (N, P), residual suspended solids, pathogenic organisms | Chemical precipitation, biological nitrification-denitrification, granular media filtration, chlorination/UV |
Preliminary and Primary Treatment
Preliminary Treatment
Preliminary treatment protects mechanical equipment, pumps, and transport conduits from mechanical blockage, abrasion, and clogging:
- Screening: Raw sewage passes through coarse, inclined bar screens followed by fine screens to intercept large floating and suspended debris (wood, rags, paper, plastic). Because these solids are predominantly inert or non-biodegradable trash, they are compacted and transferred directly to landfills or incinerators.
- Grit removal: Wastewater flows through velocity-regulated grit chambers () to allow heavy inorganic particles (sand, gravel, eggshells, broken glass) to settle by gravity while organic matter remains suspended.
Primary Treatment
Primary Sedimentation
Primary sedimentation physically removes settleable organic and inorganic suspended particulate matter:
- Screened sewage enters quiescent primary sedimentation tanks (clarifiers) with hydraulic retention times typically ranging from .
- Under gravity, suspended settleable particles consolidate at the tank bottom as primary sludge, while surface skimmers continuously remove floating scums, free oils, and grease.
Watch out. Do not conflate preliminary screening with primary sedimentation, and do not assume standard primary clarifiers always utilize chemical additives. Primary clarification functions primarily by plain gravity settling. Coagulants such as alum () are added only in chemically enhanced primary treatment (CEPT) to destabilize fine colloidal suspensions:
The insoluble gelatinous precipitates sweep fine suspended colloids out of the water column.
Secondary Biological Treatment
Secondary Treatment
Secondary treatment utilizes living microorganisms within engineered bioreactors to consume and metabolize dissolved and colloidal biodegradable organic pollutants from primary effluent. While aerobic suspended-growth systems dominate municipal wastewater engineering, anaerobic systems such as Upflow Anaerobic Sludge Blanket (UASB) reactors are effective secondary alternatives in warmer climates.
In an aerobic environment, heterotrophic bacteria oxidize carbonaceous compounds into carbon dioxide and water. Nitrogenous organic compounds (e.g., urea, ) hydrolyze to yield ammonia (), which is subsequently oxidized by autotrophic nitrifiers in two consecutive biochemical steps:
Activated Sludge Process
The activated sludge process is an aerobic suspended-growth biological treatment system configured with two interdependent units: an aeration basin and a secondary clarifier.
- Aeration and bio-flocculation: Primary effluent is mixed with an acclimated microbial culture in the aeration tank, forming the mixed liquor suspended solids (MLSS). Mechanical aerators or submerged fine-bubble diffusers continuously introduce dissolved oxygen and promote hydraulic turbulence. Heterotrophic bacteria rapidly oxidize dissolved organics, synthesizing extracellular polymeric substances (EPS) that bind bacteria, protozoa, and rotifers into settleable biological aggregates called flocs.
- Secondary clarification: The mixed liquor discharges into a quiet secondary sedimentation tank. Under quiescent conditions, biological flocs settle to the tank base, leaving a purified, clarified supernatant effluent.
- Sludge recirculation and wasting: A substantial fraction of the active settled microbial biomass is returned to the aeration basin inlet as Return Activated Sludge (RAS) to maintain a continuous, elevated biomass concentration. To balance ongoing microbial cell synthesis, excess biomass is steadily evacuated as Waste Activated Sludge (WAS) and routed to sludge stabilization.
Watch out. Biological secondary treatment converts dissolved biodegradable organic carbon into volatile , water, and settleable biomass flocs; it does not eliminate dissolved conservative inorganic salts, toxic heavy metals, or persistent non-biodegradable synthetic organics.
Tertiary Treatment, Disinfection, and Sludge Management
Tertiary Treatment
Tertiary (advanced) treatment refines water quality to meet rigorous standards, targeting residual pathogens and dissolved inorganic nutrients ( and ) that stimulate algal blooms and accelerate eutrophication:
-
Phosphorus precipitation: Addition of hydrated lime () raises the solution pH above , precipitating orthophosphate as insoluble hydroxyapatite or basic calcium phosphate.
Alternatively, coagulants such as alum () or ferric chloride () are dosed to precipitate insoluble metal phosphate complexes (, ).
-
Biological and physical nitrogen removal:
- Nitrification-Denitrification: Aerobic conversion of ammonium to nitrate (), followed by anoxic heterotrophic reduction of nitrate to inert nitrogen gas ().
- Ammonia stripping: Raising effluent pH to with lime shifts ammonium ions into dissolved ammonia gas (), which is stripped out inside counter-current packed air towers.
-
Disinfection: Polished water is treated with chlorine (, ), ozone (), or ultraviolet (UV) radiation to destroy remaining pathogens. When chlorine is added, dechlorination (using sulfur dioxide, , or sodium bisulfite, ) is performed prior to environmental release to prevent chloramine toxicity in aquatic ecosystems.
Sludge Handling and Stabilization
Solids separated from primary and secondary sedimentation consist of water and highly putrescible organic matter:
- Thickening: Gravity or dissolved air flotation tanks concentrate solids and decant free water.
- Anaerobic digestion: In heated, sealed digesters, anaerobic consortia (acidogens and methanogenic archaea) break down organic solids, producing methane-rich biogas () usable as on-site fuel.
- Dewatering: Mechanically conditioned sludge passes through plate-and-frame filter presses, belt filter presses, or centrifuges to generate a dry cake for landfilling, thermal incineration, or agricultural application.
Self-Check Questions
-
Why must a portion of the settled sludge in the secondary clarifier be continuously recycled to the aeration tank? Answer: Recycling Return Activated Sludge (RAS) sustains a concentrated population of active, acclimated microorganisms within the aeration tank. This high microbial density ensures rapid kinetic oxidation of incoming dissolved organic pollutants within a reasonable hydraulic retention time and compact reactor footprint.
-
What is the chemical role of lime ( / ) in tertiary sewage treatment? Answer: Lime supplies free calcium ions () and elevates the solution pH (). Under these alkaline conditions, dissolved soluble orthophosphate ions () precipitate out as insoluble calcium phosphate () or hydroxyapatite flocs, which are subsequently removed by sedimentation or granular filtration to prevent downstream eutrophication.
Trickling filters and the UASB process
Secondary biological sewage treatment relies on microorganisms to degrade dissolved and colloidal organic pollutants remaining after primary sedimentation. While the activated sludge process keeps microbes suspended in aerated liquid, alternative reactors immobilize biomass on solid surfaces or within dense, self-settling granules. Two widely used systems representing aerobic attached-growth and anaerobic granular-sludge technologies are the trickling filter and the Upflow Anaerobic Sludge Blanket (UASB) reactor.
Trickling filter
A trickling filter is an aerobic attached-growth biological reactor in which microorganisms grow as a slimy biofilm (zoogleal film) on a fixed packing medium. Despite its name, it does not operate by physical straining or sieving. The wide void spaces between packing pieces allow air to circulate freely by natural draft or forced ventilation while wastewater trickles downward as a thin liquid film, ensuring simultaneous access to dissolved organic substrates and atmospheric oxygen.
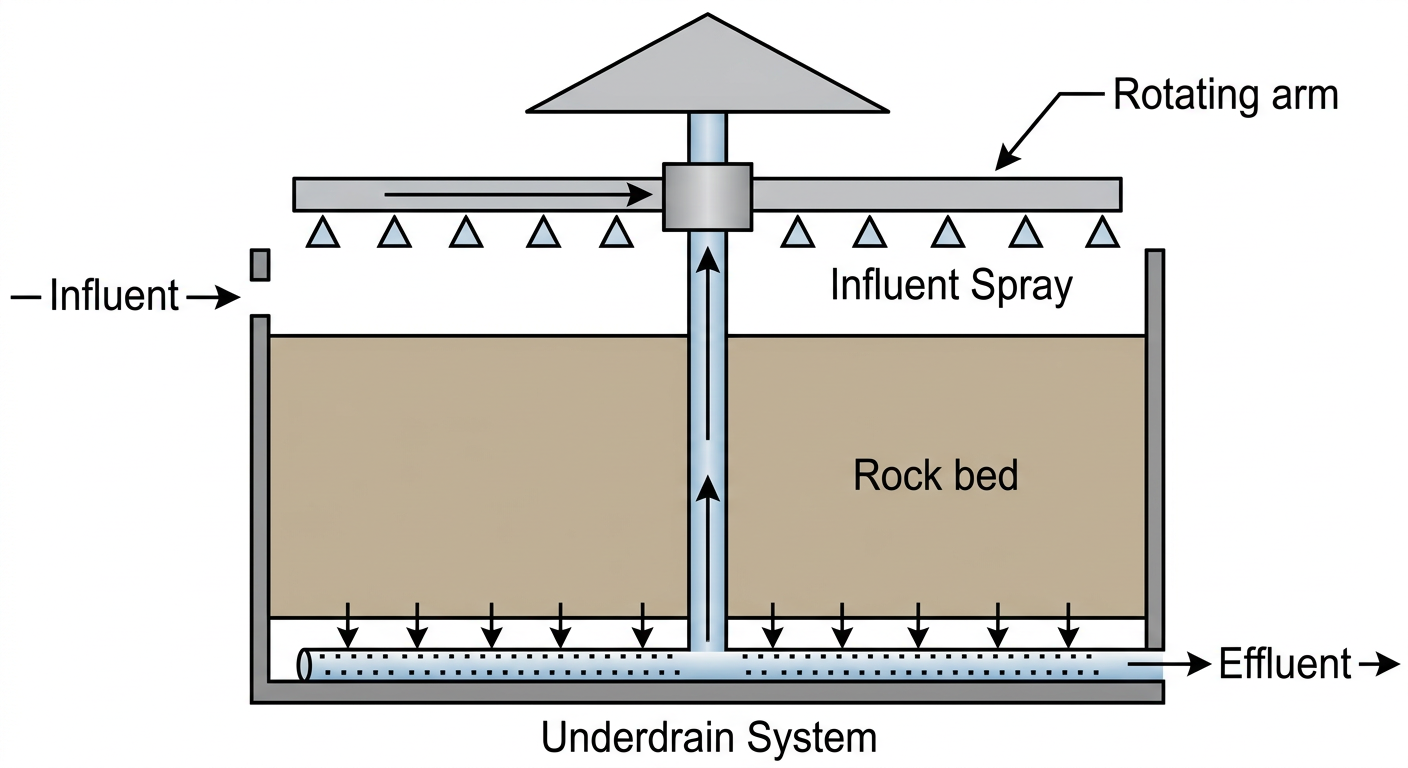
A typical trickling filter is a circular or rectangular bed packed with high-surface-area materials such as crushed rocks, broken bricks, slag, or engineered plastic modules. Treatment proceeds through the following steps:
- Distribution of pre-treated sewage: Primary clarified sewage (freed of coarse screenings and settleable grit that would otherwise clog the bed) is sprayed evenly over the top of the packing media through a rotating distributor arm.
- Biofilm contact and aerobic oxidation: As the sewage trickles downward over the media surfaces, aerobic bacteria, fungi, and protozoa within the attached biofilm absorb and oxidize dissolved organic compounds into stable end products (, , and oxidized nitrogen species such as and ).
- Sloughing of excess biomass: As the microbial layer thickens, oxygen cannot diffuse to the inner face attached to the rock or plastic. The inner microbes enter an endogenous or anaerobic state and lose their adhesion, causing patches of old biofilm to detach—a natural renewal process called sloughing.
- Underdrain collection and secondary clarification: Treated liquid and sloughed biofilm fragments exit through a perforated underdrain system at the bottom of the tank, which also functions as an air inlet. The liquid then passes into a secondary clarifier (humus tank) to settle out the sloughed biomass. Part of the clarified effluent may be recirculated to the filter inlet to keep the biofilm moist and dilute high-strength inflows.
Upflow Anaerobic Sludge Blanket process
Watch out. Student notes frequently expand UASB as "Upflow Anaerobic Sludge Bed" and claim that it treats the solid screenings removed during preliminary bar screening. Both are incorrect: UASB stands for Upflow Anaerobic Sludge Blanket, and it treats liquid wastewater (such as primary clarified sewage or high-strength industrial effluent) rather than coarse solid trash.
UASB reactor
A UASB reactor is a high-rate anaerobic biological treatment system that operates without any oxygen supply or mechanical aeration. Introducing liquid upward from the bottom at a controlled hydraulic velocity brings wastewater into continuous contact with a retained sludge blanket while washing out treated water at the top, achieving intense contact between organic pollutants and microbes without mechanical mixing.
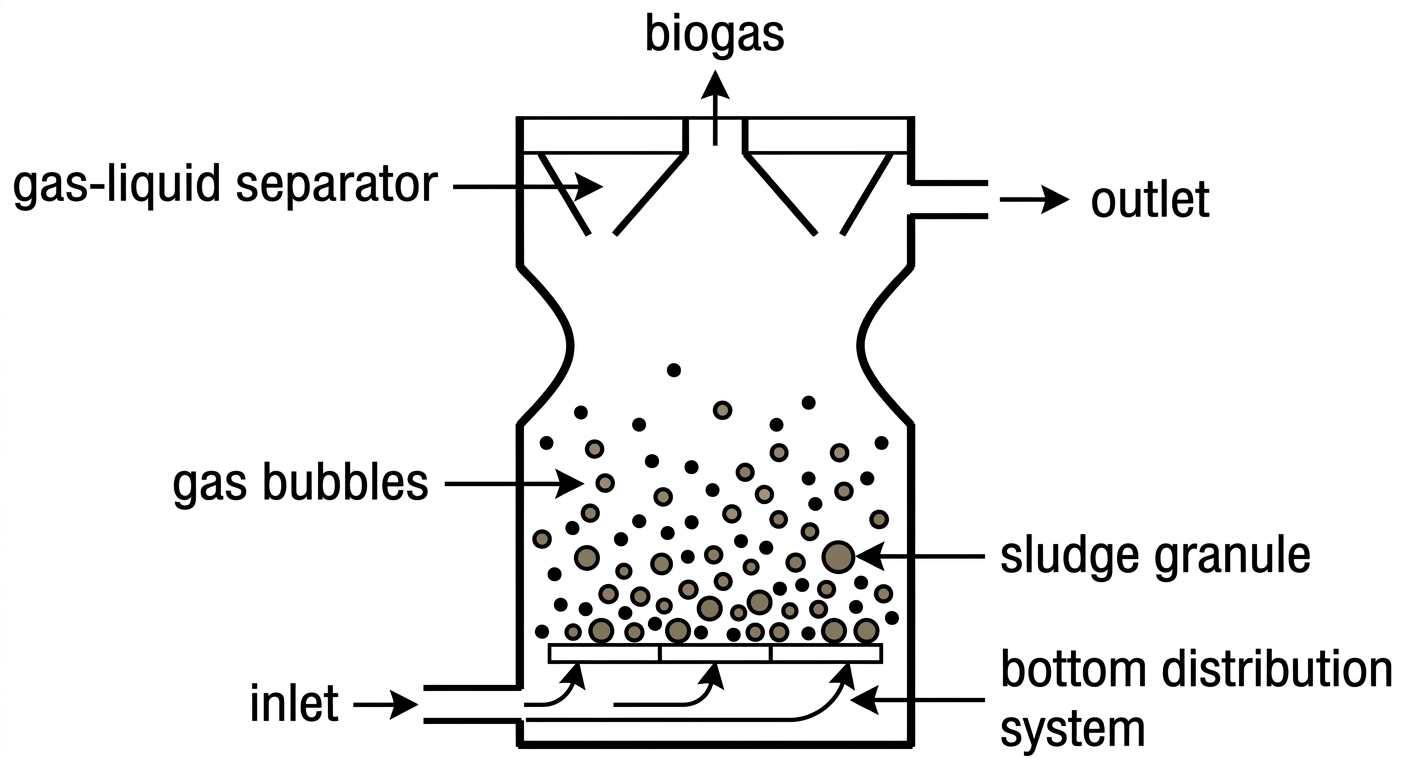
Operation of a UASB reactor involves four main features:
- Bottom feed and upward flow: Pre-treated liquid sewage enters through distribution nozzles at the bottom of the reactor and flows vertically upward. Designers select the upflow velocity according to reactor design and wastewater strength so that the sludge blanket stays suspended without being washed out.
- Granular sludge blanket: The lower zone contains a dense blanket of biologically formed sludge granules ( in diameter). These granules are compact, self-immobilized consortia of anaerobic microorganisms, including bacteria and methanogenic archaea with high settling velocities.
- Anaerobic conversion and biogas generation: As wastewater passes through the blanket, anaerobic microorganisms degrade organic matter in sequential stages—hydrolysis and acidogenesis (acid-forming bacteria converting complex organics into volatile fatty acids, , and ) followed by methanogenesis (methane-forming archaea converting intermediate acids and hydrogen into methane).
The overall process is anaerobic digestion, involving fermentation and linked oxidation–reduction reactions without free molecular oxygen. 4. Three-phase separation (Gas–Liquid–Solids Separator, GLSS): Rising biogas bubbles (primarily and , which can be captured as fuel) create natural hydraulic turbulence that mixes the blanket. At the top of the reactor, an inverted cone or gas collector dome separates the gas from liquid and deflects rising sludge granules back down into the settling zone. Clarified liquid effluent overflows from the top launder. Because anaerobic effluent still contains residual BOD, dissolved , and , it typically requires aerobic polishing and tertiary disinfection (such as chlorination) before reuse or discharge.
Comparison of trickling filter and UASB processes
| Feature | Trickling filter | UASB reactor |
|---|---|---|
| Microbial environment | Aerobic (requires atmospheric via natural draft or ventilation) | Strictly anaerobic (operates in the absence of ) |
| Biomass growth form | Biofilm attached to stationary rock or plastic media | Dense, self-granulated microbial pellets () suspended in upflow |
| Flow direction | Downflow (trickling over media from top distributor) | Upflow (bottom inlet to top gas–liquid–solids separator) |
| Energy and byproducts | Low-to-moderate energy use; produces , , and humus sludge | Can recover energy through -rich biogas fuel and very little excess sludge |
| Organic loading & BOD effect | Best suited for low-to-medium strength sewage; significantly lowers BOD via aerobic oxidation | Handles medium-to-high organic loading; lowers bulk BOD anaerobically but requires aerobic post-treatment |
| Key operational limitations | Risk of media clogging (ponding), odour, and filter flies; larger land footprint | Slow initial start-up (granule formation takes weeks/months); sensitive to cold temperatures and toxic shocks |
Self-check questions
- Why is a secondary clarifier required after a trickling filter if the microorganisms are attached to the packing media? Answer: As the biofilm grows thicker, the inner layer loses access to oxygen and detaches from the media (sloughing). A secondary clarifier is required to settle and remove this sloughed biomass from the treated effluent.
- What prevents the active bacterial biomass from washing out with the top effluent in a UASB reactor? Answer: The bacteria form dense, fast-settling granules (), and the top three-phase (gas–liquid–solids) separator deflects rising granules into a quiescent settling zone while capturing biogas in the collector dome.
E-waste and resource recovery
Nature and scope of e-waste
Electronic waste (e-waste)
Electronic waste (e-waste) refers to discarded, broken, or obsolete electrical and electronic equipment (EEE) that has reached the end of its useful life, and is discarded as waste rather than retained for continued use. Typical examples include discarded computers, mobile phones, office hardware, televisions, and household appliances such as refrigerators.
Functional second-hand products
Functional second-hand products are distinguished from e-waste by operational status and intended use. If a device still operates—or is retained for repair or consumable refilling (such as an inkjet cartridge)—and is donated or resold for its original intended purpose, it remains in active service as a reused product rather than entering the waste stream.
Material composition and component streams
E-waste is chemically and structurally complex, comprising four broad material classes: metals, plastics (such as ABS and PET), glass, and ceramics (including silica and alumina substrates). Because electronic designs evolve rapidly, exact compositions vary widely by device type and manufacturing era rather than adhering to a fixed universal ratio. E-waste represents both a rich secondary resource and a hazardous waste stream.
| Component stream | Valuable materials | Potential hazardous substances (device/era-dependent) |
|---|---|---|
| Waste printed circuit boards (WPCBs) | Precious metals () and base metals (); ceramic and polymer substrates | Lead () in legacy solders; brominated flame retardants in structural resins |
| Display units (CRTs, LCDs, LEDs) | Copper deflection yokes, aluminium structural frames, glass, engineering plastics () | Lead in cathode ray tube (CRT) funnel glass; mercury () in cold-cathode fluorescent lamps (CCFLs) used in older liquid crystal displays (LCDs) |
| Batteries | Cobalt, lithium, nickel, lead, copper | Lead in lead-acid cells; cadmium () in cells; reactive electrolytes in lithium-ion cells |
| Capacitors and specialized parts | Aluminium, tantalum, copper leads | Polychlorinated biphenyls (PCBs) in legacy dielectric fluids; beryllium oxide () ceramics in high-power RF components |
Watch out. Do not confuse the two meanings of PCB in e-waste chemistry: printed circuit boards are the laminated fiberglass-resin substrates supporting electronic components, whereas polychlorinated biphenyls (not "bisphenyls") are toxic, persistent chlorinated hydrocarbons historically used as dielectric insulating fluids in transformers and capacitors. Additionally, note that LCD denotes liquid crystal display (not "diode"), and while older CCFL-backlit LCD panels contain mercury, modern light-emitting diode (LED) backlights do not contain mercury lamps.
Environmental pathways and occupational health risks
When e-waste is deposited in unlined landfills, openly incinerated, or processed through informal primitive recycling (such as open-wire burning or crude acid baths), hazardous toxins migrate through multiple environmental compartments:
- Atmospheric release: Open burning and uncontrolled smelting release volatile metal fumes (), fine particulate matter, and halogenated dioxins/furans generated from brominated flame retardants.
- Soil and groundwater contamination: Acidic rainwater percolating through unlined disposal sites leaches bioavailable heavy metals and hexavalent chromium () directly into surrounding soils and underlying aquifers.
- Occupational exposure: Workers operating without localized dust extraction or specialized personal protective equipment (PPE) face direct toxic exposure via inhalation of metal fumes, absorption of volatile species, and ingestion of settled particulates.
| Hazardous substance | Primary biological and health impacts |
|---|---|
| Lead () | Neurotoxicant; impairs the central and peripheral nervous systems, inhibits heme synthesis, compromises renal function, and causes severe irreversible developmental neurotoxicity in children. |
| Cadmium () | Nephrotoxicant with a long biological half-life; bioaccumulates in the renal cortex and liver, causing chronic kidney failure, osteomalacia, and bone fragility. |
| Mercury () | Bioaccumulative neurotoxin; causes chronic damage to the central nervous system and kidneys, accompanied by severe cognitive, sensory, and respiratory dysfunctions. |
| Beryllium () | Group 1 human carcinogen; chronic inhalation of particulate dust or fumes during mechanical crushing and milling induces chronic beryllium disease (berylliosis), an incurable granulomatous lung disorder. |
4R management hierarchy and resource recovery
Sustainable handling of post-consumer electronics follows the Reduce, Reuse, Recycle, and Recover hierarchy, supported by integrated industrial infrastructure:
-
Reduce:
- Minimize waste generation at source by preventing unnecessary technological upgrades.
- Design durable, modular hardware with extended operating lifetimes and select multi-functional electronic devices.
-
Reuse:
- Prolong the working life of functional electronics via secondary resale markets, institutional donation programs, modular hardware refurbishment, and component re-manufacturing (e.g., refilling printer cartridges).
-
Recycle (collection, dismantling, and pre-processing):
- Formal collection and segregation: Systematic diversion of e-waste from standard municipal solid waste streams prevents mechanical compaction and accidental breakage of toxic components.
- Manual dismantling and depollution: Certified technicians systematically isolate and extract hazardous sub-assemblies—such as batteries, mercury CCFL tubes, leaded CRT funnel glass, and legacy dielectric capacitors—prior to bulk mechanical processing.
- Size reduction and physical sorting: Depolluted equipment hulks are crushed and shredded into uniform fragments, followed by automated multi-stage physical separation (magnetic sorting for ferrous metals, eddy-current separation for non-ferrous metals like and , and density-based air classification for polymers).
-
Recover (metallurgical refining):
- Resource extraction: High-grade target streams (primarily concentrated WPCBs) undergo industrial pyrometallurgical smelting, hydrometallurgical selective leaching, or electrometallurgical refining within facilities outfitted with acid gas scrubbers and closed-loop wastewater treatment.
- Circular economy value: "Urban mining" of copper, gold (), and silver () from e-waste yields dramatically higher metal grades than natural ores, drastically lowering the net carbon and energy footprints of primary extraction. This circular loop advances key United Nations Sustainable Development Goals: SDG 12 (Responsible Consumption and Production), SDG 6 (Clean Water and Sanitation, via leachate prevention), and SDG 3 (Good Health and Well-Being).
Self-check questions
-
Why must dismantling and depollution occur before e-waste is mechanically shredded? Answer: Shredding intact e-waste would rupture batteries, mercury lamps, and leaded glass, contaminating the recyclable plastic and metal streams and exposing workers to toxic dust and vapours.
-
Why is urban mining of waste printed circuit boards environmentally preferable to mining virgin ore? Answer: WPCBs contain significantly higher concentrations of copper, gold, and silver than natural ores, so well-managed recovery can reduce virgin material demand and mining impacts while keeping hazardous materials out of uncontrolled disposal. Energy benefits depend on the feed and recovery process.
Greenhouse gases and climate change
Weather
Weather refers to short-term atmospheric conditions (temperature, humidity, atmospheric pressure, wind, and precipitation) occurring over hours or days in a localized region.
Climate
Climate is the statistical average and variability of weather patterns over extended periods, typically decades or centuries (conventionally 30 years).
Global warming
Global warming is the sustained, long-term increase in Earth's global mean surface temperature, primarily driven by anthropogenic emissions.
Climate change
Climate change encompasses the broader spectrum of long-term shifts in planetary climate systems, including shifts in temperature patterns, altered precipitation regimes, and increased frequency of extreme weather events, driven by natural cycles or human activities.
Mechanism of the greenhouse effect
The Earth's equilibrium surface temperature is dictated by the radiative balance between incoming shortwave solar energy and outgoing terrestrial thermal heat:
- Solar transmission and surface absorption: Incoming solar radiation consists primarily of short-wavelength radiation (ultraviolet and visible light). A fraction is reflected by clouds and atmospheric aerosols or absorbed directly, while the majority passes through the atmosphere and is absorbed by the planetary land and ocean surfaces, warming them.
- Terrestrial longwave emission: The warmed Earth re-radiates thermal energy back toward space at much longer wavelengths, primarily in the thermal infrared (IR) spectrum ().
- Molecular absorption and re-emission: Greenhouse gases (GHGs) present in the troposphere absorb specific bands of this outgoing terrestrial IR radiation. These excited molecules re-emit thermal radiation isotropically (in all directions)—directing a substantial fraction back down toward the planetary surface and lower troposphere, thereby trapping thermal energy.
Watch out. Greenhouse gases do not primarily warm the planet by directly trapping incoming ultraviolet (UV) or shortwave solar rays on their way in. Instead, they absorb outgoing longwave thermal infrared radiation emitted by the warmed Earth.
Infrared activity and molecular symmetry
To absorb infrared radiation via the greenhouse mechanism, a molecule must possess infrared-active vibrational modes—meaning that its electric dipole moment must fluctuate dynamically as its chemical bonds stretch or bend.
| Gas Classification | Chemical Examples | Molecular Symmetry & Dipole Behavior | IR Spectral Activity |
|---|---|---|---|
| Inactive Bulk Gases | , , | Homonuclear diatomic or monatomic; highly symmetric charge distributions; stretching produces no dipole moment change (). | IR transparent; cannot absorb terrestrial thermal radiation. |
| Active Greenhouse Gases | , , , , , Halocarbons (CFCs, HFCs) | Polyatomic or heteronuclear; asymmetric bond stretches and angle-bending vibrational modes induce a fluctuating dipole moment (). | IR active; strongly absorb and re-emit outgoing thermal IR photons. |
Natural versus enhanced greenhouse effect
Natural greenhouse effect
The natural greenhouse effect is a fundamental planetary process essential for sustaining life. Without baseline levels of naturally occurring water vapour and carbon dioxide, Earth's mean surface equilibrium temperature would be approximately (well below the freezing point of water) rather than the current hospitable average of .
Enhanced greenhouse effect
The enhanced (anthropogenic) greenhouse effect is the additional radiative forcing caused by the rapid accumulation of long-lived greenhouse gases in the atmosphere since the onset of the Industrial Revolution, trapping excess thermal energy and destabilizing global climate equilibria.
Water vapour feedback
Water vapour () is quantitatively the most abundant greenhouse gas. However, its residence time in the atmosphere is extremely short (days to weeks), and its atmospheric concentration is governed strictly by the Clausius-Clapeyron thermodynamic relationship (evaporation vs. condensation as a function of temperature). Consequently, water vapour acts as an amplifying feedback loop—initial warming induced by non-condensable GHGs increases tropospheric moisture capacity, which traps more heat—rather than a direct anthropogenic forcing agent.
Anthropogenic vs. natural emission sources
Human-driven greenhouse gas additions are categorized across four principal sectors:
-
Fossil fuel combustion: The extraction and oxidation of coal, petroleum, and natural gas for electricity, heavy industry, heating, and transportation releases fossil carbon stored for millions of years:
-
Deforestation and land-use change: Rapid land clearing and burning permanently eliminates terrestrial photosynthetic carbon sinks while converting stored biomass carbon into atmospheric .
-
Agriculture and livestock management:
- Enteric fermentation in ruminant livestock and anaerobic decomposition in flooded rice paddies generate biogenic methane ().
- Excessive application of synthetic nitrogen fertilizers accelerates microbial nitrification and denitrification pathways, releasing potent nitrous oxide ().
-
Industrial chemical processes: Thermal calcination of limestone during cement manufacturing directly liberates carbon dioxide:
Additionally, specialized manufacturing, refrigeration, semiconductor etching, and foam blowing emit synthetic fluorinated gases (CFCs, HFCs, ).
Watch out. Volcanic eruptions are a natural geological source of and aerosols, not an anthropogenic (human-caused) driver of climate change.
Environmental and engineering consequences
- Hydrological disruption and thermal extremes: Elevated atmospheric temperatures accelerate evaporation rates in arid zones, exacerbating prolonged agricultural droughts and soil degradation. Concurrently, warm air holds more water vapour (approximately per rise), leading to hyper-concentrated precipitation events, storm surges, and severe urban flash flooding.
- Sea-level rise: Global mean sea levels are driven upward by two physical phenomena:
- Thermal expansion: Liquid water expands volumetrically as ocean heat content rises.
- Land-ice ablation: The mass melting of terrestrial glaciers, continental ice fields, and the Greenland and Antarctic ice sheets discharges new liquid volume into ocean basins. (Note: The melting of floating Arctic sea ice does not directly alter sea level because floating ice already displaces its equivalent mass in water, though its disappearance lowers planetary surface albedo, accelerating regional warming.)
- Ecological and biochemical stress: Marine heatwaves trigger widespread coral bleaching and reduce the saturation solubility of dissolved oxygen () in surface waters, stressing aquatic life. Shifting climatic zones also modify the geographical ranges of disease vectors (e.g., mosquitoes transmitting malaria and dengue).
(Note: While CFCs act as greenhouse gases in the troposphere, stratospheric ozone depletion itself is a distinct photochemical issue concerning UV radiation rather than the principal driver of global warming; this is examined in the next section.)
Mitigation versus adaptation
Engineering solutions to climate destabilization are organized into proactive source-reduction (mitigation) and defensive vulnerability-reduction (adaptation).
| Evaluation Parameter | Climate Mitigation | Climate Adaptation |
|---|---|---|
| Core Objective | Eliminate or reduce greenhouse gas emissions at the source and enhance planetary carbon sinks to limit total warming. | Re-engineer human infrastructure, systems, and ecological management to withstand unavoidable impacts. |
| Timescale & Scope | Long-term, global benefits (because well-mixed atmospheric GHGs distribute uniformly around the globe). | Immediate to near-term, local and regional vulnerability focus. |
| Engineering Interventions | • Transition to renewable energy (solar, wind, nuclear, hydro)<br>• Carbon Capture and Storage (CCS) at point sources<br>• Fleet electrification and grid modernization<br>• Industrial waste-heat recovery and energy optimization<br>• Large-scale afforestation and soil carbon sequestration | • Construction of coastal seawalls, dykes, and tidal surge barriers<br>• Upgraded, high-capacity urban stormwater drainage networks<br>• Developing genetically engineered drought- and salt-tolerant crops<br>• Heat-resilient civil materials and passive urban cooling architecture |
Self-check questions
-
Question: Why is an active greenhouse gas whereas , which makes up most of the atmosphere, is not? Answer: is a linear triatomic molecule possessing asymmetric stretching and bending vibrational modes. These vibrations alter its molecular dipole moment during motion, enabling it to absorb and emit photons in the thermal infrared spectrum. In contrast, is a symmetric homonuclear diatomic molecule whose stretching vibration produces no change in dipole moment (), rendering it transparent to thermal infrared radiation.
-
Question: Why is atmospheric water vapour classified as a climate feedback rather than a primary anthropogenic forcing agent? Answer: The atmospheric residence time of water vapour is brief (on the order of days), and its concentration is governed thermodynamically by ambient atmospheric temperature (through evaporation-condensation equilibria) rather than direct industrial discharge rates. When long-lived anthropogenic forcing agents (such as and ) warm the troposphere, the warmer air accommodates a higher partial pressure of water vapour, which subsequently absorbs additional infrared radiation and amplifies the baseline warming trend.
Stratospheric ozone and ozone depletion
Ozone
Ozone () plays two opposing environmental roles depending on its atmospheric altitude:
- Tropospheric ozone: In the lower atmosphere, ground-level ozone is a secondary air pollutant and respiratory irritant formed via photochemical smog reactions.
- Stratospheric ozone: Located primarily in the ozone layer between above Earth's surface, it functions as a vital protective radiation shield by absorbing biologically harmful solar ultraviolet (UV-B and UV-C) radiation before it reaches the biosphere.
Natural formation and the Chapman cycle
Chapman cycle
The Chapman cycle describes the continuous photostationary steady state through which stratospheric ozone is naturally generated and decomposed:
-
Photodissociation of molecular oxygen: High-energy short-wave UV photons () cleave molecular oxygen into reactive oxygen free radicals:
-
Ozone formation: An oxygen radical combines with molecular oxygen in the presence of an inert collision partner (typically or ). The third body dissipates excess vibrational energy so that the newly formed bond does not immediately rupture:
-
Photodissociation of ozone: Ozone absorbs solar UV-B radiation, preventing it from reaching the surface while regenerating and an oxygen radical (which can rapidly reform via Step 2, converting UV radiation into thermal energy):
-
Natural recombination: An oxygen radical collides with an ozone molecule, permanently destroying both odd-oxygen species:
Watch out. Do not write ozone formation as . The combination of and is exothermic and requires a third-body collision partner to carry away excess energy. Solar UV radiation drives the reverse photodissociation reaction.
Catalytic ozone depletion
Ozone depletion
Ozone depletion is the anthropogenic thinning of the stratospheric ozone layer caused by chemical catalysts accelerating recombination reactions far beyond natural rates.
The primary historical agents are chlorofluorocarbons (CFCs, such as or Freon-12), previously used as refrigerants, aerosol propellants, and blowing agents. Because CFCs are chemically inert and water-insoluble in the troposphere, they migrate upward into the stratosphere. There, intense short-wavelength UV radiation cleaves the carbon-chlorine bond via photolysis to liberate reactive chlorine radicals:
The chlorine radical acts as a homogeneous catalyst in an efficient chain reaction:
Net catalytic reaction:
Because is regenerated at the end of each cycle, a single chlorine radical can destroy upwards of ozone molecules before being trapped in stable reservoir species (such as or ). In polar spring, heterogeneous reactions on polar stratospheric clouds (PSCs) release active chlorine, causing intense seasonal ozone depletion (the "ozone hole") via dimer cycles that bypass the requirement for free atoms.
Watch out. In the second step of the standard gas-phase chlorine cycle, reacts with an oxygen radical (), not with another molecule. Writing is a common error.
Depletion by nitrogen oxides ()
Nitric oxide radicals ()—emitted directly into the stratosphere by high-altitude supersonic aircraft or formed via the oxidation of tropospheric nitrous oxide ()—catalyse an analogous destruction cycle:
Net reaction:
Environmental impacts and mitigation
Thinning of the stratospheric ozone layer allows elevated levels of biologically harmful UV-B radiation () to penetrate to ground level:
- Human health: Induces pyrimidine dimer formation in cellular DNA, causing elevated rates of non-melanoma and melanoma skin cancers, cataracts, and immune suppression.
- Terrestrial and marine ecosystems: Inhibits photosynthesis and biomass production in terrestrial crops; damages phytoplankton populations at the foundation of marine food webs.
Montreal Protocol
The Montreal Protocol on Substances that Deplete the Ozone Layer (1987) mandated the global phase-out of CFCs, halons, and carbon tetrachloride. These substances were initially replaced with transitional hydrochlorofluorocarbons (HCFCs) and subsequently with non-ozone-depleting hydrofluorocarbons (HFCs) and hydrofluoroolefins (HFOs), allowing gradual recovery of the stratospheric ozone shield.
Distinguishing ozone depletion from the greenhouse effect
Stratospheric ozone depletion and the enhanced greenhouse effect are fundamentally distinct physical phenomena:
| Feature | Stratospheric Ozone Depletion | Enhanced Greenhouse Effect |
|---|---|---|
| Primary radiation involved | Incoming solar short-wave ultraviolet (UV-B) | Outgoing terrestrial long-wave infrared (IR) |
| Atmospheric region | Stratosphere () | Troposphere () |
| Underlying mechanism | Free-radical catalytic destruction of molecules | Absorption and re-emission of terrestrial thermal IR radiation |
| Key anthropogenic agents | CFCs, halons, stratospheric | , , , fluorinated greenhouse gases |
| Direct primary consequence | Increased UV-B surface flux (skin cancer, cataracts, ecological damage) | Tropospheric warming, altered climate patterns, thermal ocean expansion |
Self-check questions
-
Why can a small concentration of stratospheric destroy a vastly larger molar quantity of ? Answer: Photolysis of releases a radical that oxidises to form , which subsequently reacts with an oxygen atom to regenerate . Because the halogen radical acts as a homogeneous catalyst, a single radical survives long enough to repeat the catalytic cycle thousands of times before terminating.
-
What is the role of in the reaction ? Answer: is a neutral third-body collision partner (such as or ) that carries away the excess vibrational energy released during the exothermic formation of ozone. Without , the nascent molecule would retain sufficient energy to spontaneously dissociate back into reactants.
Sustainable development and the 17 SDGs
Principles of Sustainable Development
Sustainable development
Sustainable development, as defined by the Brundtland Commission, is development that meets the needs of the present without compromising the ability of future generations to meet their own needs. It is built on the principle of intergenerational fairness: current resource consumption, industrial growth, and waste generation must not degrade the ecological systems that future populations will depend upon.
Achieving sustainability requires integrating environmental protection, social equity, and economic viability into long-term planning:
- An industrial process that maximizes short-term profit by discharging untreated effluent shifts remediation, public health, and ecological restoration costs onto society and future generations.
- Engineering projects must balance immediate utility with environmental resilience and lifecycle impact.
Watch out. Sustainable development does not mean infinite, unconstrained economic growth, nor does it mean halting all industrial activity. Because Earth has finite resources and limited capacity to assimilate waste, sustainable engineering focuses on long-term resource efficiency, circular material flows, and staying within ecological boundaries.
The Three Pillars of Sustainability
Sustainable decision-making balances three interconnected pillars. A project is truly sustainable only where all three overlap.
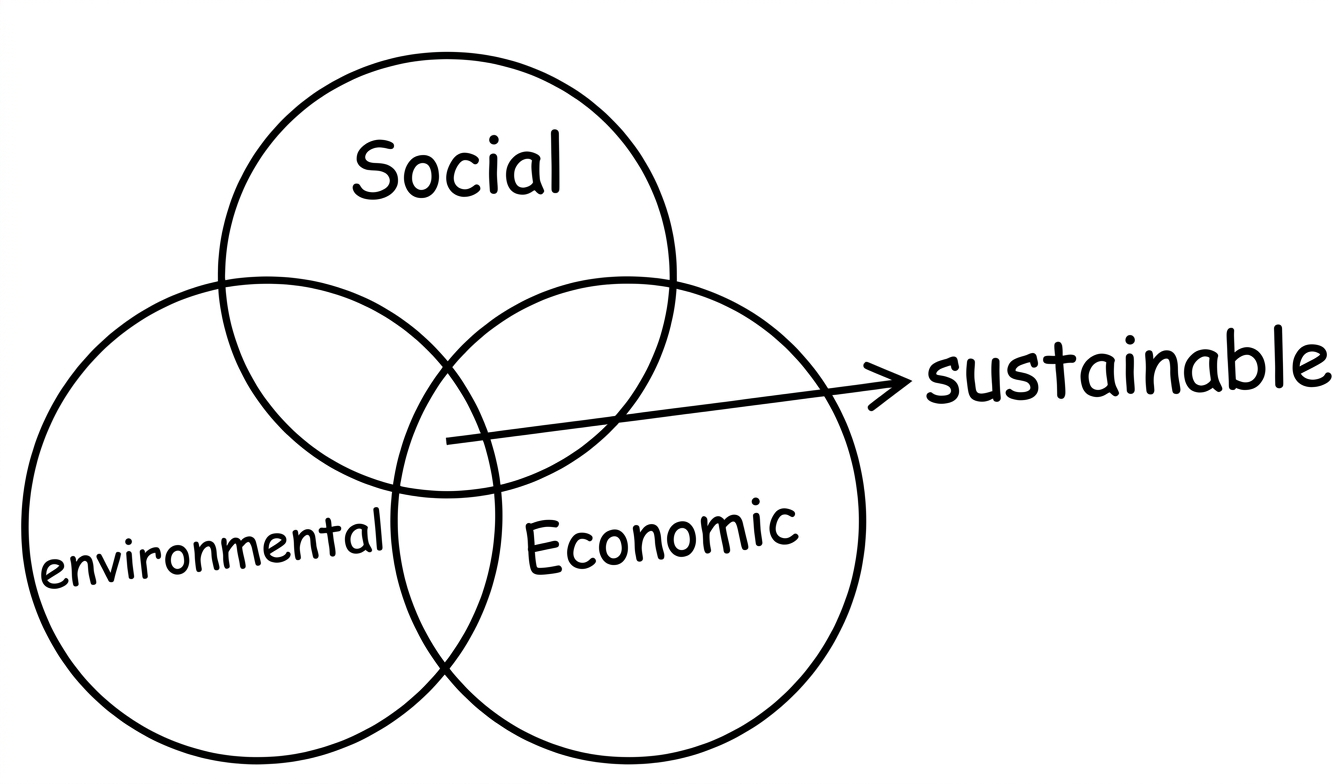
Figure: The three overlapping pillars of sustainable development—Environmental, Social, and Economic—intersecting to define sustainable practice.
| Pillar | Core Objective | Engineering Application (Water & Materials) |
|---|---|---|
| Environmental protection | Preserve ecosystems, biodiversity, and natural resources through pollution control and waste minimization. | Treating sewage to lower BOD before discharge; recycling e-waste metals; implementing water reuse and zero-liquid-discharge (ZLD) systems. |
| Social inclusion | Ensure equitable access to resources, health, safety, and opportunities across all demographics and generations. | Providing safe, pathogen-free drinking water to all communities; preventing toxic e-waste exposure among informal workers. |
| Economic prosperity | Maintain efficient, productive systems that deliver long-term value without externalizing environmental or social harm. | Designing treatment plants for lifecycle affordability, energy recovery (such as UASB biogas), and manageable maintenance costs. |
The 17 Sustainable Development Goals (SDGs)
In 2015, the United Nations adopted the 2030 Agenda for Sustainable Development, establishing 17 interconnected global goals to address poverty, inequality, environmental degradation, and peace:
- No Poverty
- Zero Hunger
- Good Health and Well-being
- Quality Education
- Gender Equality
- Clean Water and Sanitation
- Affordable and Clean Energy
- Decent Work and Economic Growth
- Industry, Innovation and Infrastructure
- Reduced Inequalities
- Sustainable Cities and Communities
- Responsible Consumption and Production
- Climate Action
- Life Below Water
- Life on Land
- Peace, Justice and Strong Institutions
- Partnerships for the Goals
Alignment of Environmental Chemistry with the SDGs
The chemical and treatment processes studied across this module directly support several global goals:
- SDG 6 (Clean Water and Sanitation): Supported by water purification—coagulation, filtration, and disinfection (chlorination, ozonation, UV)—alongside biological sewage treatment (activated sludge, trickling filters, UASB) and desalination. Note that water softening (ion exchange) primarily protects industrial boilers and piping from scale; safe drinking water requires pathogen disinfection and toxic contaminant removal, not softening alone.
- SDG 3 (Good Health and Well-being): Disinfecting drinking water prevents waterborne diseases, while proper e-waste handling prevents heavy-metal poisoning from lead, mercury, and cadmium.
- SDG 12 (Responsible Consumption and Production): Hydrometallurgical and pyrometallurgical recovery of precious and base metals from e-waste reduces virgin ore mining and hazardous landfilling.
- SDG 13 (Climate Action): Mitigating greenhouse gas emissions (, , , CFCs) and capturing methane from anaerobic digesters (UASB) for fuel directly limit global warming.
- SDG 7 (Affordable and Clean Energy): Transitioning from fossil fuels to renewable energy and recovering biogas from wastewater sludge improve energy sustainability.
- SDG 14 (Life Below Water) & SDG 15 (Life on Land): Removing organic load (BOD/COD) and nutrients (nitrates and phosphates) during tertiary sewage treatment prevents eutrophication and hypoxia in aquatic ecosystems, while controlling acid rain and hazardous waste protects terrestrial habitats.
Engineering Trade-Offs in Sustainable Design
No engineering technology offers an unlimited environmental benefit; every solution involves measurable trade-offs across the three pillars.
Watch out. Do not treat pollution-control or water-treatment technologies as having zero environmental cost. For example, reverse osmosis (RO) produces low-salinity permeate from brackish or saline sources; drinking-water use also requires appropriate post-treatment (supporting SDG 6), but high-pressure pumps consume significant electrical energy (challenging SDG 7 and SDG 13) and generate concentrated reject brine that can harm marine life if discharged improperly (challenging SDG 14).
Lifecycle thinking
Lifecycle thinking is the evaluation and quantification of energy demand, chemical consumption, by-product disposal, and social affordability before selecting an engineering process. For instance, breakpoint chlorination provides reliable residual disinfection, yet reaction with natural organic matter can form carcinogenic trihalomethanes (THMs). In contrast, ozonation and UV avoid chlorinated by-products but require higher capital and electrical input and leave no protective residual in distribution pipes.
Self-check questions
-
Question: How does the Brundtland definition of sustainable development connect current engineering choices to future generations?
Answer: It requires that present needs be met without compromising the ability of future generations to meet their own needs, meaning engineers must prevent irreversible resource depletion and long-term environmental pollution. -
Question: Give one example of an engineering trade-off between SDG 6 (Clean Water and Sanitation) and SDG 13 (Climate Action) or SDG 14 (Life Below Water) in seawater desalination.
Answer: Reverse osmosis desalination supplies clean water (SDG 6), but requires high electrical energy—often increasing greenhouse gas emissions (SDG 13)—and produces concentrated brine that can damage ecosystems without suitable concentrate management (SDG 14).
Revision sheet and practice problems
Core Formulas and Concept Sheet
Water Hardness and Equivalents
Calcium Carbonate Standard
Calcium carbonate (, molar mass , equivalent mass ) serves as the universal reference standard because it is a familiar sparingly soluble scale-forming compound and its molecular mass of greatly simplifies stoichiometric calculations.
To convert any hardness-producing ion or dissolved salt into equivalent (or ) as :
For solutions of known normality (where ):
| Species | Molar Mass () | Equivalent Mass () | Species | Molar Mass () | Equivalent Mass () |
|---|---|---|---|---|---|
Hardness Partitioning and Ion-Balance Rules
Total Hardness ()
Total Hardness is the combined sum of equivalents of all multivalent cations—principally calcium () and magnesium (). Monovalent ions () do not consume soap and are excluded.
Carbonate / Temporary Hardness ()
Carbonate Hardness is the hardness associated with bicarbonate and carbonate anions. It is strictly bounded by both total hardness and total alkalinity ():
Non-Carbonate / Permanent Hardness ()
Non-Carbonate Hardness is the hardness caused by sulfates, chlorides, or nitrates that cannot be removed by boiling:
Watch out. Adding does not change total, temporary, or permanent hardness because cannot precipitate soap. Adding increases alkalinity, which can convert permanent hardness into temporary hardness, but Carbonate Hardness can never exceed Total Hardness ( is never negative).
Unit Conversions and Oxygen Demand
-
Hardness Units:
- (in dilute aqueous solution)
- (1 grain per Imperial gallon)
-
Unseeded : For initial dissolved oxygen , 5-day dissolved oxygen , and dilution factor :
Quick Retrieval Questions
-
What happens when bicarbonate hardness is heated?
Calcium bicarbonate precipitates as calcium carbonate (). Prolonged heating of magnesium bicarbonate can ultimately form magnesium hydroxide (). Removal depends on operational temperature and duration and is not perfectly quantitative. -
How are the two demineralization beds regenerated?
The hydrogen-form cation exchange resin () is regenerated with a strong acid ( or ). The hydroxide-form anion exchange resin () is regenerated with sodium hydroxide (). Sodium-cycle zeolite softening differs fundamentally: it replaces and with , retaining dissolved salts. -
What drives reverse osmosis?
An applied external hydrostatic pressure on the feed side that exceeds the osmotic pressure difference across the semi-permeable membrane, providing the required net driving force for pure water permeation. -
What does breakpoint chlorination signify?
It marks the point where readily oxidizable species and ammonia-related combined chlorine residuals have been fully oxidized and destroyed. Dosing past this point produces a directly proportional, stable free-chlorine residual. It does not imply the complete oxidation of all refractory organic matter. -
Why use dechlorination?
Residual chlorine can form toxic disinfection byproducts or harm downstream aquatic ecosystems. Reducing agents such as sodium sulfite (), sulfur dioxide (), or granular activated carbon (GAC) beds neutralize free chlorine before discharge. -
Why is COD usually higher than BOD, and can one be calculated from the other?
uses a boiling, acidified strong dichromate oxidant that chemically breaks down both biodegradable and non-biodegradable refractory organics, whereas measures only biologically assimilable carbon over 5 days. There is no universal theoretical conversion factor between the two. -
How do sewage-treatment stages differ?
- Preliminary: Bar screens and grit chambers remove coarse debris and dense mineral grit to protect mechanical pumps.
- Primary: Quiescent clarifiers allow settleable suspended organic solids to settle as raw sludge.
- Secondary: Aerobic biological reactors (e.g., activated sludge) digest dissolved organics into settleable biomass flocs.
- Advanced (Tertiary): Nutrient removal (nitrogen and phosphorus), sand filtration, or disinfection for reuse.
-
How does UASB differ from activated sludge?
The Upflow Anaerobic Sludge Blanket (UASB) process operates anaerobically without supplemental aeration, cultivating dense granular sludge with methanogenic archaea that convert organics into methane-rich biogas. Activated sludge is an aerobic process requiring continuous mechanical aeration, secondary clarification, and high rates of activated sludge recycling. -
Why must e-waste be managed separately?
Electronic waste combines valuable, recoverable resources (gold, copper, rare earth elements) with hazardous heavy metals (lead, cadmium, mercury) and brominated flame retardants. Uncontrolled landfilling or open burning releases toxic persistent pollutants into groundwater and the atmosphere. -
How do greenhouse warming and ozone depletion differ?
Greenhouse warming is caused by greenhouse gases () trapping terrestrial infrared radiation in the troposphere. Ozone depletion occurs in the stratosphere, where UV-C radiation photolyzes chlorofluorocarbons (), releasing chlorine free radicals that catalytically destroy the stratospheric ozone layer. -
What connects this module to sustainable development?
Effective water purification and wastewater treatment directly support SDG 6 (Clean Water and Sanitation); metal recovery from e-waste and industrial wastes supports SDG 12 (Responsible Consumption and Production); and mitigating greenhouse gas emissions addresses SDG 13 (Climate Action).
Numerical Practice Problems
-
Single-salt hardness:
Calculate the hardness in as of a water sample containing of .Solution:
The molar mass of is .
Since valence , the equivalent mass is . -
Ion-balance hardness:
A water sample contains , , and . Calculate total, carbonate (temporary), and non-carbonate (permanent) hardness.Solution:
Convert each concentration to equivalents using respective equivalent masses (, , ):Applying the ion-balance partition:
-
Alkalinity cap and non-hardness salts:
A water sample has a total hardness of as and total bicarbonate alkalinity of as (due to dissolved ), along with of . Determine , , and .Solution:
- contains monovalent ions and contributes to hardness.
- Carbonate hardness cannot exceed total hardness regardless of excess sodium bicarbonate alkalinity:
-
Molarity to hardness:
Calculate the hardness of a solution in as .Solution:
Each molecule yields one ion ().
Normality . -
Five-day BOD calculation:
An unseeded wastewater sample diluted to ( dilution) has an initial dissolved oxygen of , which drops to after of incubation at . Calculate .Solution:
Substitute , , and dilution factor :
Long-Answer Review Prompts
- Explain why is chosen as the reference standard for water hardness, and derive the relationship between temporary, permanent, and total hardness when sodium bicarbonate is present.
- With neat chemical equations, describe the complete demineralization of hard water using cation and anion exchange resins and explain the regeneration step.
- Define reverse osmosis, explain how it desalinates brackish or seawater, and state its operational advantages.
- Sketch and explain the breakpoint chlorination curve, detailing the chemical reactions occurring in each zone and how excess chlorine is removed.
- Compare Dissolved Oxygen (DO), Biochemical Oxygen Demand (BOD), and Chemical Oxygen Demand (COD) in terms of definition, measurement time, and significance for aquatic life.
- Describe the preliminary, primary, secondary and advanced stages of municipal sewage treatment, distinguishing ordinary settling from chemically enhanced clarification.
- Compare aerobic and anaerobic oxidation of organic sewage, and explain the working of a UASB reactor.
- Discuss the photochemical equilibrium of stratospheric ozone, the free-radical mechanism of -induced ozone depletion, its biological consequences, and mitigation measures.
Final Learning Checklist
- Convert dissolved salts and ions into equivalents and apply without producing negative permanent hardness.
- Write balanced chemical equations for soap scum formation, boiling of bicarbonates, ion-exchange softening/regeneration, alum coagulation, and dechlorination.
- Distinguish between ion-exchange demineralization, reverse osmosis, breakpoint chlorination, ozonation, and UV disinfection.
- Calculate and explain the process flows of activated sludge, trickling filters, and UASB reactors.
- Summarize e-waste hazards/recycling, greenhouse warming, stratospheric ozone chemistry, and the UN SDGs.